FDA CDRH issues guidance on notifications of medical device shortages during COVID-19

The US Food and Drug Administration (FDA) Center for Devices and Radiological Health (CDRH) published a guidance describing how medical device manufacturers should notify CDRH of shortages during the COVID-19 emergency.
This is a new reporting requirement for medical device manufacturers, as they are generally not required to report shortages. The notification requirement will continue for the duration of the COVID-19 emergency.
Keypoints of the guidance:
- Manufacturers of devices critical to the public health such as ventilators, hemodialysis equipment, automatic external defibrillators, cardiopulmonary bypass oxygenators, specimen collection kits, reagents for serological testing, and pulse oximeters and other monitoring equipment.
- CDRH must be notified at least six months in advance of a permanent discontinuance or interruption in manufacturing of a device that is likely to lead to a meaningful disruption in supply in the US. If that timeline is not realistic, that manufacturer should notify CDRH as soon as possible and not later than 7 days after the discontinuance or interruption in manufacturing.
- Permanent discontinuance or interruption in manufacturing, as well as the reasons for such discontinuance or interruption must be notified to the CDRH. Manufacturers should also include appropriate identifying information, including marketing submission holder name, marketing submission number, manufacturer name, FDA establishment identifier, device name, product code and contact information.
- Notification should be submitted via email to the CDRH. An initial notification may include a list of all affected devices. If an additional device is affected after the initial notification, and new notification should be submitted to the FDA.
For more details, please see the guideline from FDA.
Meditrial can help – Contact us for immediate support
Meditrial has developed rapid workflows to conduct risk assessment, implement rescue plans and execute solutions for ongoing clinical trials:
- Identify patients requiring follow-up
- Conduct investigator and study nurse calls for training to address the situation
- Obtain authorizations for decentralized data collection
- Support to patients for alternative access to needed care
- Prepare procedures and worksheets for remote collection of patient clinical data
- Monitor work instructions and trainings for site support and remote monitoring
- EDC and ePRO solutions to enable study continuity
- Privacy protection measures
Contact Meditrial for immediate assistance in Europe or the US.
Table of Contents
Getting a CE mark for medical devices: a step-by-step guide
Practical guidance on the CE mark for medical devices under EU MDR, from classification and evidence to conformity assessment and EUDAMED
A Practical RBQM Framework for Medical Device Trials
Risk-based quality management for medical device trials, with practical actions for risk assessment, monitoring, data review, and trial oversight
EU Breakthrough Medical Devices Under MDCG 2025-9 (BtX)
EU breakthrough medical devices gain a clearer regulatory pathway under MDCG 2025-9, with early advice and lifecycle evidence planning
What Swissmedic Fast-Track Means for Clinical Trial Sponsors
Swissmedic swissethics Fast-Track Pilot Project accelerates eligible trial reviews and makes coordinated regulatory preparation increasingly important.
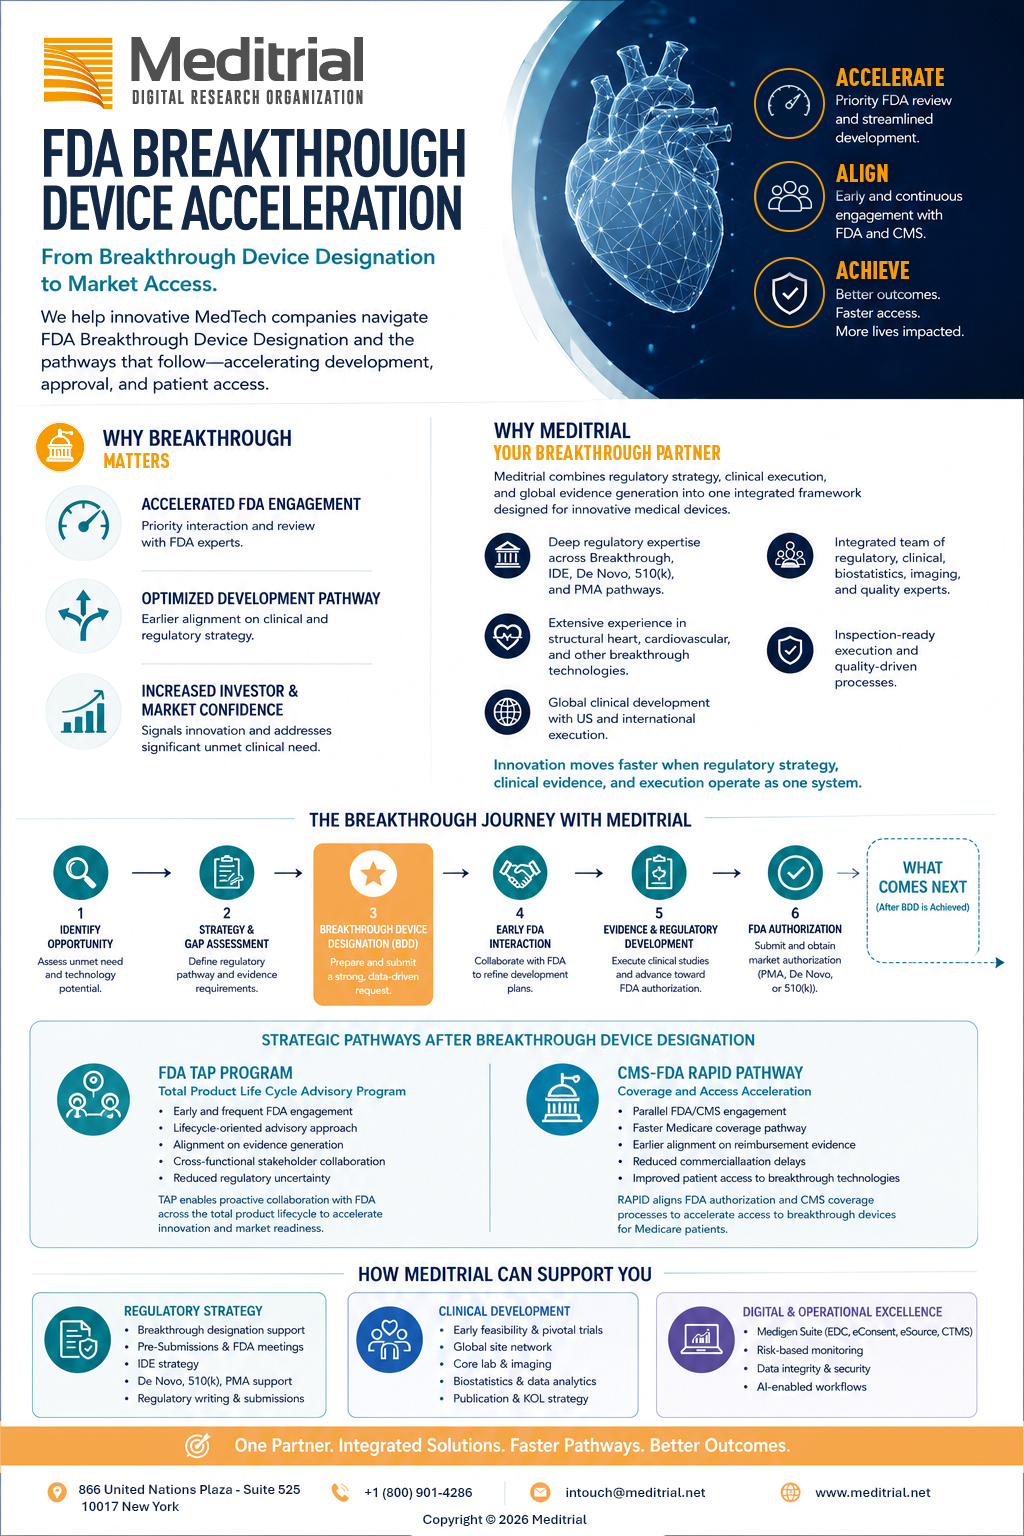
FDA Breakthrough Device Acceleration
How innovative MedTech companies can accelerate FDA approval, secure Medicare coverage, and reach patients faster—starting with Breakthrough Device Designation.
The contribution of patient associations in clinical trials
Historically, patient organizations have had the primary purpose of supporting the communities they represented. Today, this remains a crucial function, but modes have changed, as has the role that associations and patients themselves take on [...]
This article provides general information and does not constitute regulatory or legal advice. Requirements should be confirmed for the specific study, applicable regulatory framework, and markets concerned.