Germany: BfArM and PEI issued supplementary recommendations on management of clinical trials during COVID-19 pandemic

BfArM (Bundesinstitut für Arzneimittel und Medizinprodukte) and PEI (Paul-Ehrlich-Institut) provided supplementary recommendations to the guideline on management of clinical trials during COVID-19 released by the European authorities on 20th March 2020.
Key points of BfArM and PEI recommendations on remote source data verification procedures (rSDV) in Germany:
- rSDV should only be considered in exceptional cases and only to the extent necessary.
- rSDV requires written approval by the investigator and the site. In Germany approval by the regulatory authority and ethics committee is required as well. In the application the planned rSDV approach needs to be described, starting with a study-specific risk assessment. This risk assessment can be part of the study protocol (Quality Control and Quality Assurance section) or an appendix to the protocol.
- SOPs and study-specific documents (those on risk-based quality management and monitoring) need to be adapted accordingly.
- The method chosen for rSDV (the EMA guidelines lists three options) depends on the infrastructure of the sites. In the application for approval, the sponsor shall list all rSDV methods which may apply because only the ones submitted can obtain approval.
- For each rSDV method, the source document/data made available under rSDV, as well as information on precautions taken to preserve confidentiality of data and privacy of trials subjects must be provided.
- The guidance then lists specific information required for each of the 3 rSDV approaches. It also covers requirements for document management systems and telemedicine. Additional data protection considerations may apply.
For more details, please see the official communication from the Paul-Ehrlich-Institut.
Meditrial can help – Contact us for immediate support
Meditrial has developed rapid workflows to conduct risk assessment, implement rescue plans and execute solutions for ongoing clinical trials:
- Identify patients requiring follow-up
- Conduct investigator and study nurse calls for training to address the situation
- Obtain authorizations for decentralized data collection
- Support to patients for alternative access to needed care
- Prepare procedures and worksheets for remote collection of patient clinical data
- Monitor work instructions and trainings for site support and remote monitoring
- EDC and ePRO solutions to enable study continuity
- Privacy protection measures
Contact Meditrial for immediate assistance in Europe or the US.
Table of Contents
Getting a CE mark for medical devices: a step-by-step guide
Practical guidance on the CE mark for medical devices under EU MDR, from classification and evidence to conformity assessment and EUDAMED
A Practical RBQM Framework for Medical Device Trials
Risk-based quality management for medical device trials, with practical actions for risk assessment, monitoring, data review, and trial oversight
EU Breakthrough Medical Devices Under MDCG 2025-9 (BtX)
EU breakthrough medical devices gain a clearer regulatory pathway under MDCG 2025-9, with early advice and lifecycle evidence planning
What Swissmedic Fast-Track Means for Clinical Trial Sponsors
Swissmedic swissethics Fast-Track Pilot Project accelerates eligible trial reviews and makes coordinated regulatory preparation increasingly important.
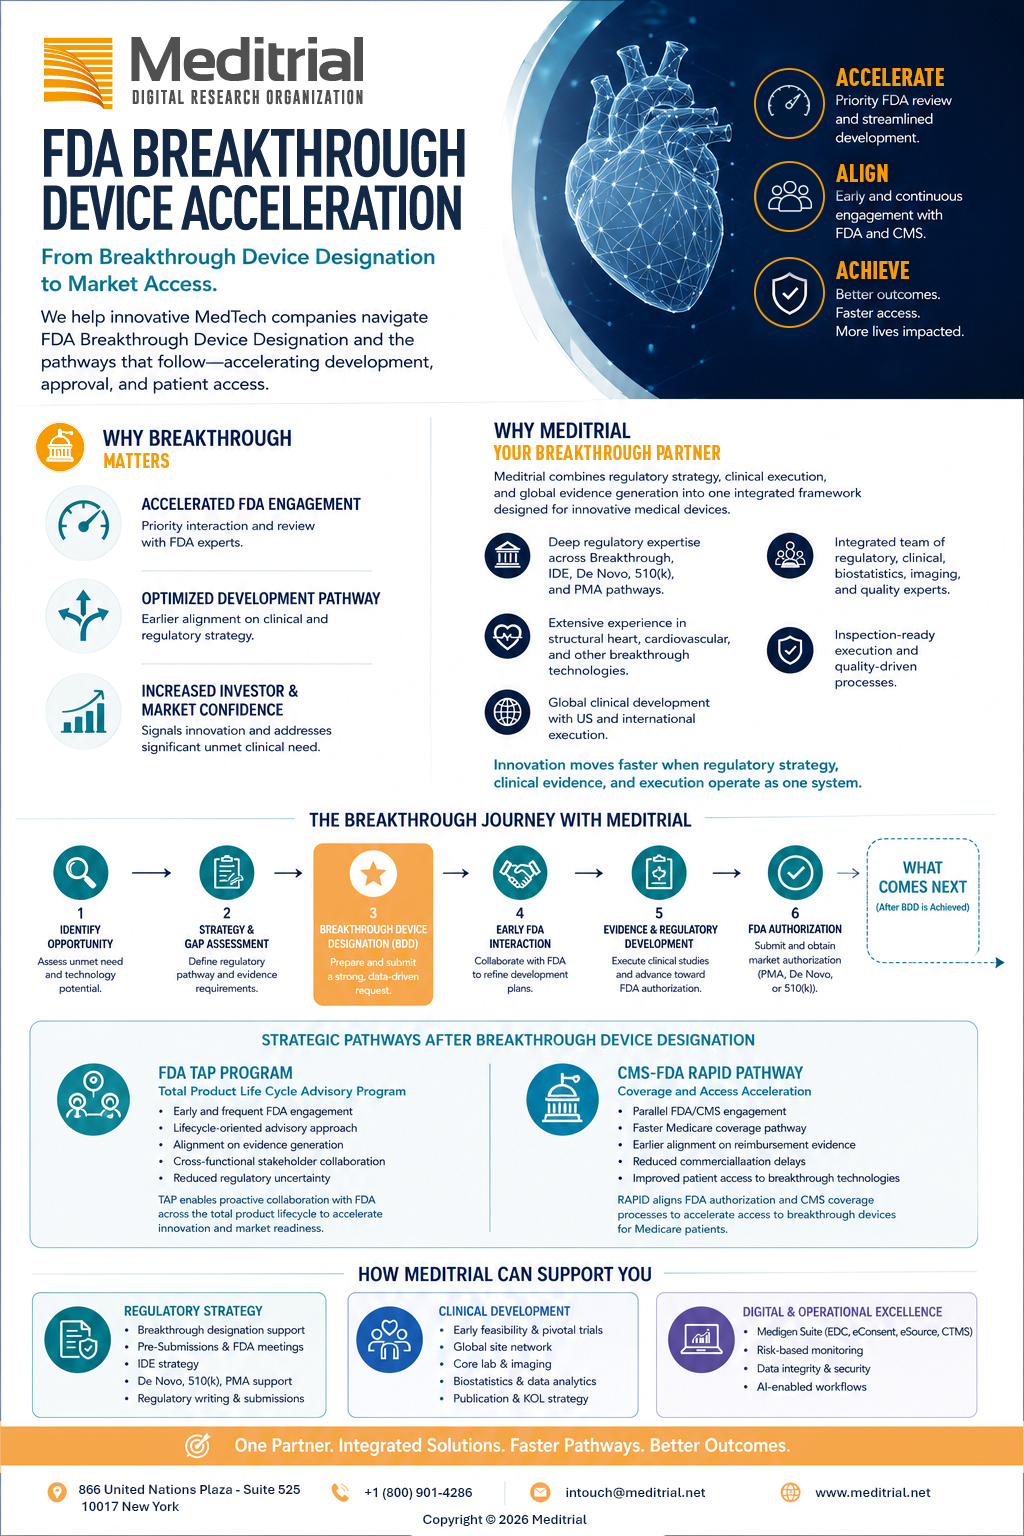
FDA Breakthrough Device Acceleration
How innovative MedTech companies can accelerate FDA approval, secure Medicare coverage, and reach patients faster—starting with Breakthrough Device Designation.
The contribution of patient associations in clinical trials
Historically, patient organizations have had the primary purpose of supporting the communities they represented. Today, this remains a crucial function, but modes have changed, as has the role that associations and patients themselves take on [...]
This article provides general information and does not constitute regulatory or legal advice. Requirements should be confirmed for the specific study, applicable regulatory framework, and markets concerned.