US: FDA published finalized guidance on inspections of medical device establishments

The Food and Drug Administration (FDA) issued guidance on inspections of medical device establishments. The finalized document follows draft guidance issued on March 28, 2019, filling a requirement of the FDA Reauthorization Act of 2017 (FDARA).
The guidance specifies how the Agency will implement uniform processes and standards that are related to inspections of foreign and domestic device establishments. It also describes standardized methods of communication during the inspection process and identifies practices for investigators and device establishments to enable the continuity of inspections of such establishments.
Pre-announcement Notice and Communication
FDARA indicates a ‘reasonable time’ within which the pre-announcement of inspections of device establishment should take place. National inspections will be pre-announced no less than 5 calendar days in advance of the inspection, while pre-announcement of foreign inspection will generally take place more than 5 calendar days before the inspection.
Standard Inspection Timeframe
FDA standards for reasonably estimated timeframes of inspections generally range from 3 to 6 continuous business days and are based on the type of surveillance inspection and the extent of covered required for a pre-approval review. Additionally, it may be necessary to extend the duration of an inspection for a “number of reasons”.
Communication During Inspections
FDA states in its guidance that regular verbal communications during the inspection about its status between the investigator and the owner, operator, or agent in charge of the device establishment should be maintained.
For more details, please see the FDA guidance.
Meditrial has developed rapid workflows to conduct risk assessment, implement rescue plans and execute solutions for ongoing clinical trials:
- Identify patients requiring follow-up
- Conduct investigator and study nurse calls for training to address the situation
- Obtain authorizations for decentralized data collection
- Support to patients for alternative access to needed care
- Prepare procedures and worksheets for remote collection of patient clinical data
- Monitor work instructions and trainings for site support and remote monitoring
- EDC and ePRO solutions to enable study continuity
- Privacy protection measures
Contact Meditrial for immediate assistance in Europe or the US.
Table of Contents
Getting a CE mark for medical devices: a step-by-step guide
Practical guidance on the CE mark for medical devices under EU MDR, from classification and evidence to conformity assessment and EUDAMED
A Practical RBQM Framework for Medical Device Trials
Risk-based quality management for medical device trials, with practical actions for risk assessment, monitoring, data review, and trial oversight
EU Breakthrough Medical Devices Under MDCG 2025-9 (BtX)
EU breakthrough medical devices gain a clearer regulatory pathway under MDCG 2025-9, with early advice and lifecycle evidence planning
What Swissmedic Fast-Track Means for Clinical Trial Sponsors
Swissmedic swissethics Fast-Track Pilot Project accelerates eligible trial reviews and makes coordinated regulatory preparation increasingly important.
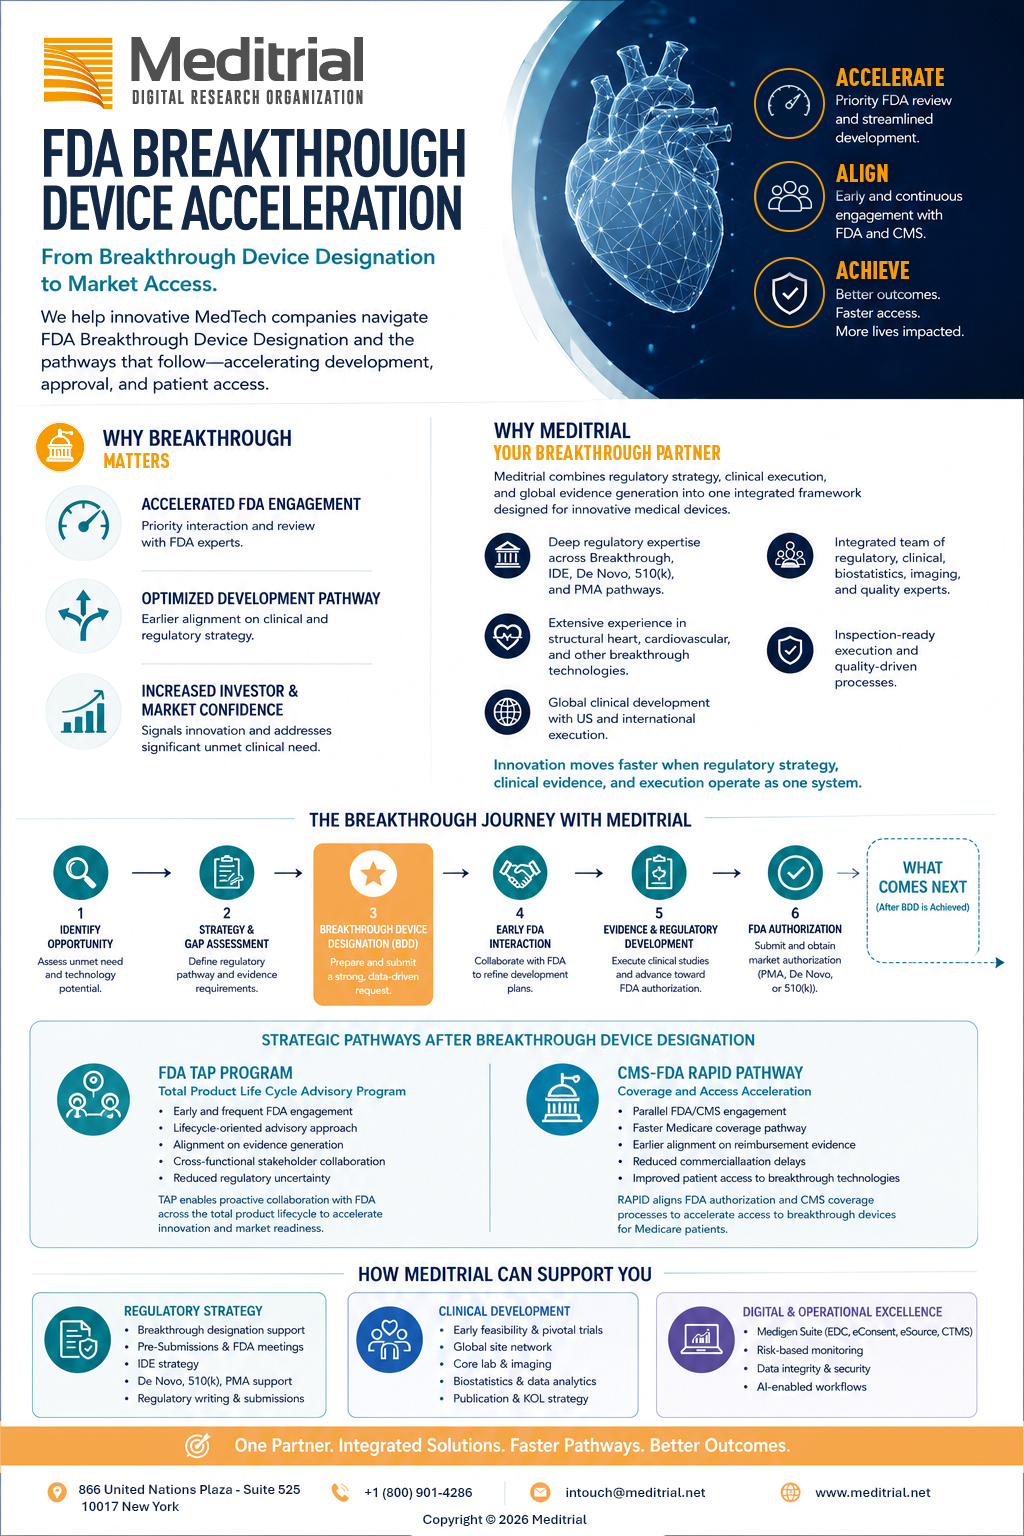
FDA Breakthrough Device Acceleration
How innovative MedTech companies can accelerate FDA approval, secure Medicare coverage, and reach patients faster—starting with Breakthrough Device Designation.
The contribution of patient associations in clinical trials
Historically, patient organizations have had the primary purpose of supporting the communities they represented. Today, this remains a crucial function, but modes have changed, as has the role that associations and patients themselves take on [...]
This article provides general information and does not constitute regulatory or legal advice. Requirements should be confirmed for the specific study, applicable regulatory framework, and markets concerned.