EU: Medical Device Single Audit Program guidance by the EU Commission’s Medical Device Coordination Group
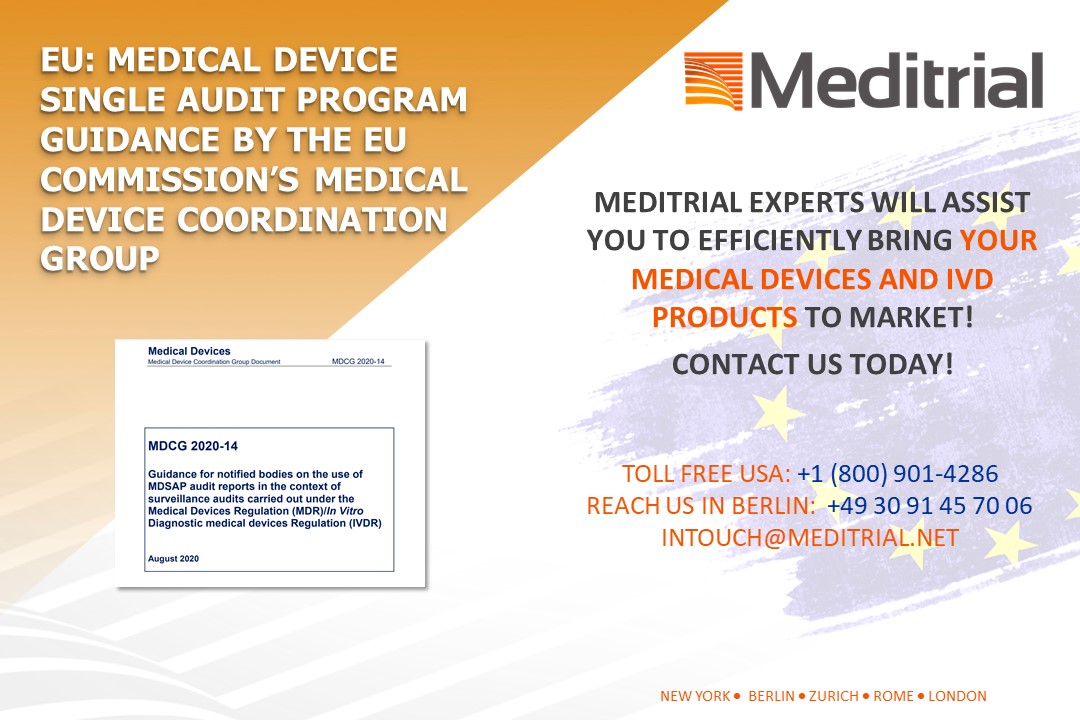

The EU Commission’s Medical Device Coordination Group issued a guidance providing details on how notified bodies can reference Medical Device Single Audit Program audit reports when planning surveillance audits under the MDR or IVDR. Meditrial can help you to get your medical or in-vitro device documentation ready for the next audit by EU or US authorithies. Reach out to Meditrial today!
Guidance keypoints:
- MDSAP audit reports cannot replace the annual surveillance audits required under MDR/IVDR
- MDSAP audit reports within the EU legislative framework are possible only where the MDSAP audit covers similar or equivalent MDR/IVDR requirements
- the scope and outputs of manufacturers’ recent MDSAP audit reports can be taken into account as an input for developing surveillance audit programmes
- nonconformities raised in MDSAP audit reports can trigger the notified body to pay particular attention to those aspects in the MDR/IVDR planned surveillance audit
- MDSAP audit reports should not be used for initial quality management system audits required for EU QMS certificates
- Reports from unannounced MDSAP audits or special audits, should not be taken into account in narrowing of focus in MDR/IVDR surveillance audits.
For more details, please see the guidance from the Medical Device Coordination Group.
Contact Meditrial for immediate assistance in Europe or the US.
Table of Contents
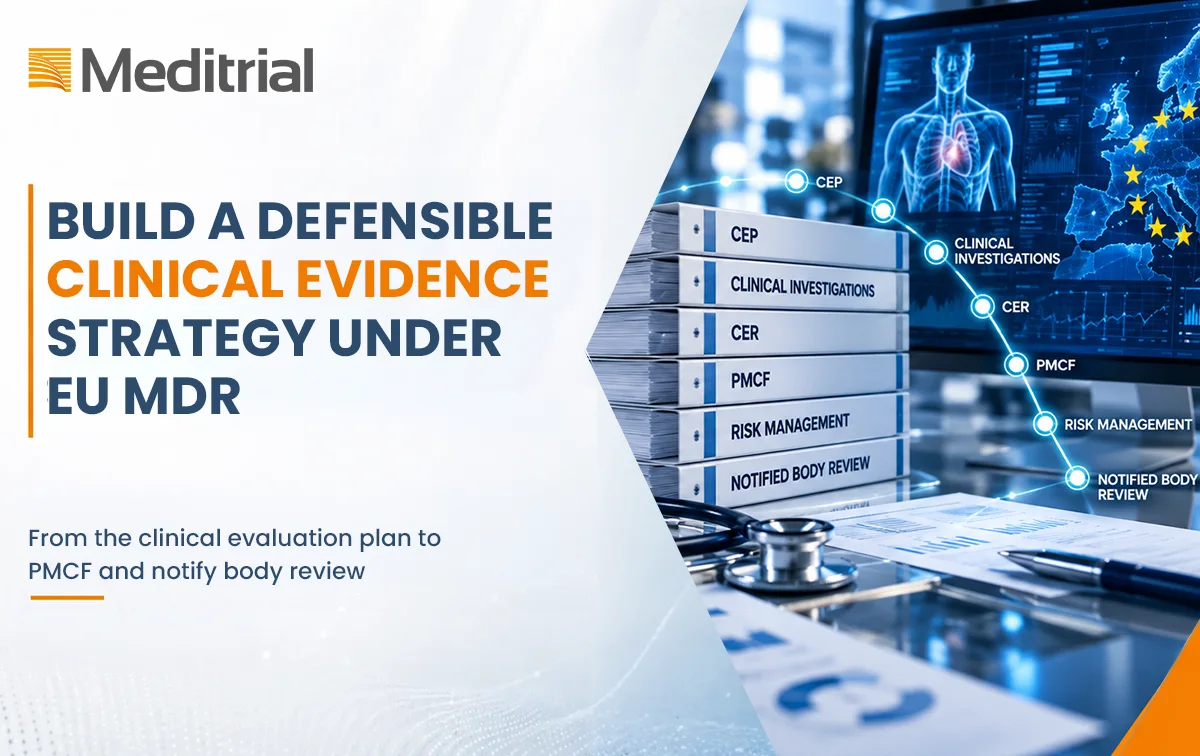
Build a Defensible Clinical Evidence Strategy Under EU MDR
Build a defensible clinical evidence strategy under MDR Annex XIV, from the clinical evaluation plan to PMCF and notified body review.
Getting a CE mark for medical devices: a step-by-step guide
Practical guidance on the CE mark for medical devices under EU MDR, from classification and evidence to conformity assessment and EUDAMED
A Practical RBQM Framework for Medical Device Trials
Risk-based quality management for medical device trials, with practical actions for risk assessment, monitoring, data review, and trial oversight
EU Breakthrough Medical Devices Under MDCG 2025-9 (BtX)
EU breakthrough medical devices gain a clearer regulatory pathway under MDCG 2025-9, with early advice and lifecycle evidence planning
What Swissmedic Fast-Track Means for Clinical Trial Sponsors
Swissmedic swissethics Fast-Track Pilot Project accelerates eligible trial reviews and makes coordinated regulatory preparation increasingly important.
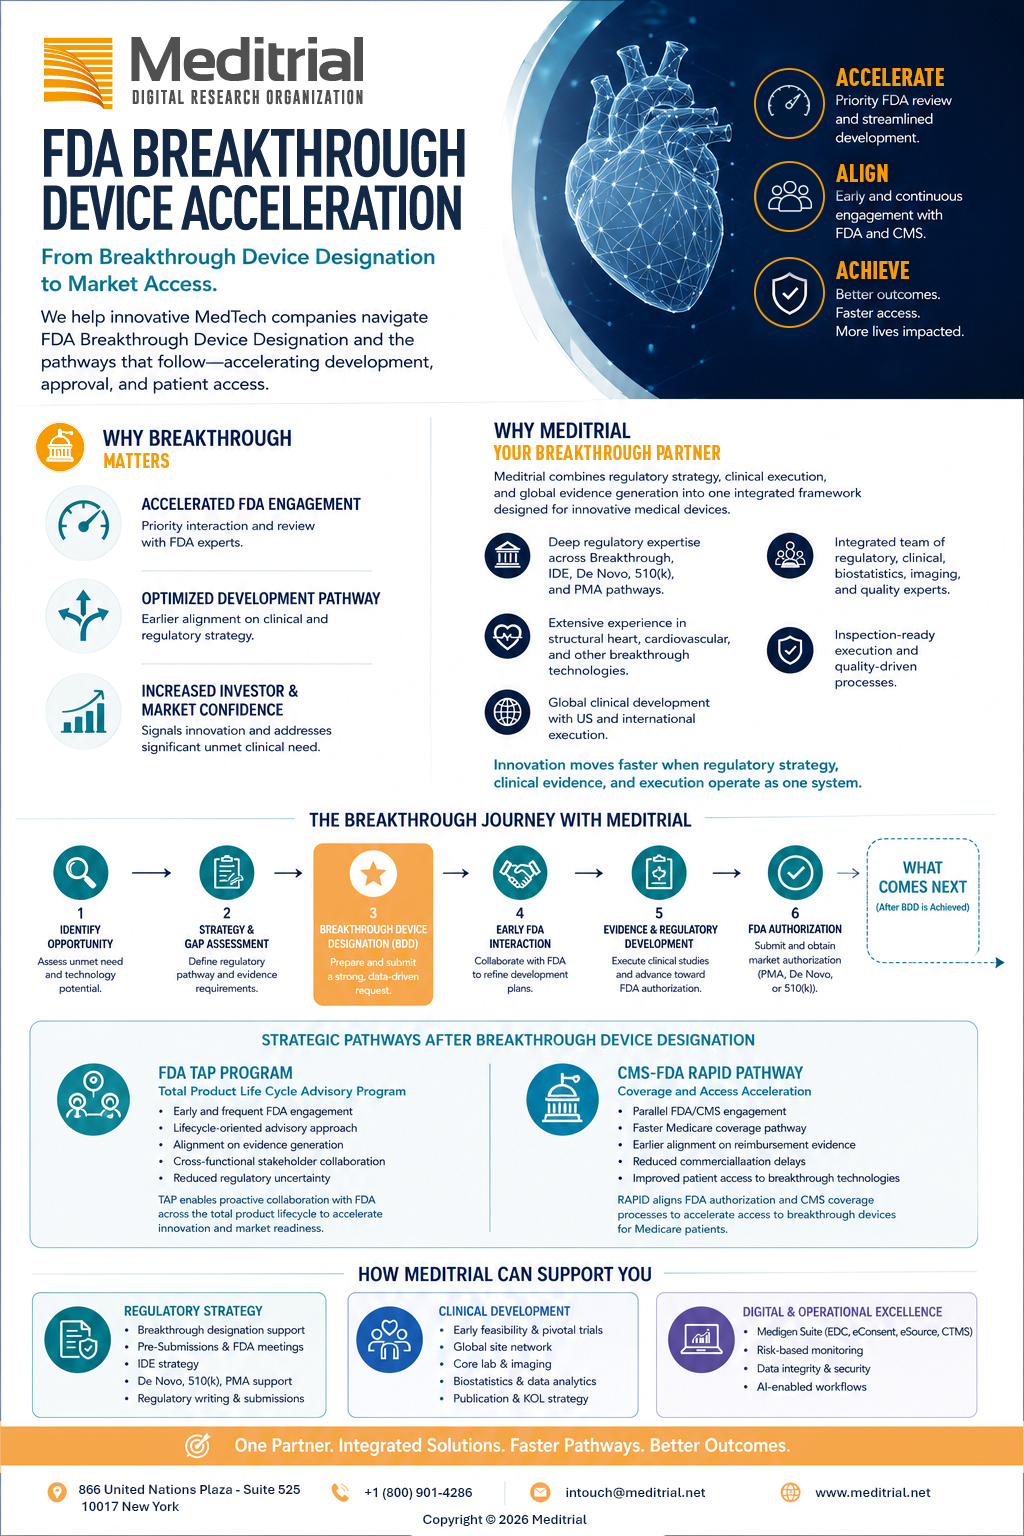
FDA Breakthrough Device Acceleration
How innovative MedTech companies can accelerate FDA approval, secure Medicare coverage, and reach patients faster—starting with Breakthrough Device Designation.
This article provides general information and does not constitute regulatory or legal advice. Requirements should be confirmed for the specific study, applicable regulatory framework, and markets concerned.