New Guidance: MDCG 2022-3 Verification by Notified Bodies
In accordance with the conformity assessment procedures (described in Article 48 of IVDR), in addition to quality management system and technical documentation assessments, notified bodies are required to verify product batches of class D IVDs in line with Annexes IX and XI of the IVDR. Therefore, in order to verify the conformity of class D IVDs, manufacturers should forward to notified bodies the reports on tests carried out on each manufactured batch of class D devices IVDR. Moreover, the manufacturer should make available samples of manufactured class D devices or batches of devices to the notified body in accordance with pre-agreed conditions and detailed arrangements.
Meditrial provides global regulatory strategy and approval services
Receive dedicated support in every stage of the technology lifecycle approval, maximizing compliance, performing correct tests.
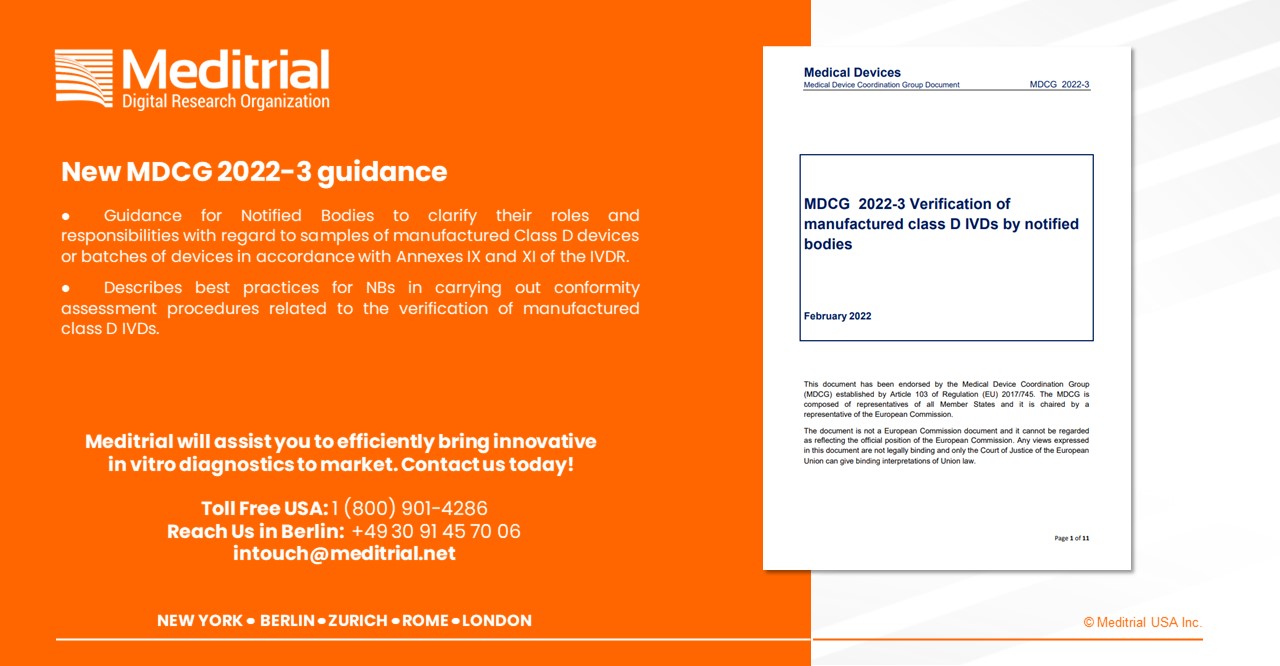
The newly published guidance MDCG 2022-3 Verification of manufactured class D IVDs by notified bodies aims to provide guidance to notified bodies to clarify their role and responsibilities in relation to samples of manufactured class D devices or batches of devices in accordance with Annexes IX and XI of the IVDR.
This guidance also describes best practices for notified bodies in performing conformity assessment procedures relating to the verification of manufactured class D IVDs.
It also includes guidance on:
- relevant information to be included within the notified body’s procedures,
- the content of the required pre-agreed conditions and detailed arrangements between the notified body and the manufacturer (IVDR Annex IX Section 4.12 & Annex XI Section 5.1),
- guidance on the frequency of samples of devices or batches of devices to be sent to the EURL (IVDR Annex IX Section 4.12 & Annex XI Section 5.1)
Read more in the MDCG guidance.
Meditrial’s medical, regulatory and technical experts will assist you to efficiently bring innovative in vitro diagnostics to market. Contact us today!
Table of Contents
Getting a CE mark for medical devices: a step-by-step guide
Practical guidance on the CE mark for medical devices under EU MDR, from classification and evidence to conformity assessment and EUDAMED
A Practical RBQM Framework for Medical Device Trials
Risk-based quality management for medical device trials, with practical actions for risk assessment, monitoring, data review, and trial oversight
EU Breakthrough Medical Devices Under MDCG 2025-9 (BtX)
EU breakthrough medical devices gain a clearer regulatory pathway under MDCG 2025-9, with early advice and lifecycle evidence planning
What Swissmedic Fast-Track Means for Clinical Trial Sponsors
Swissmedic swissethics Fast-Track Pilot Project accelerates eligible trial reviews and makes coordinated regulatory preparation increasingly important.
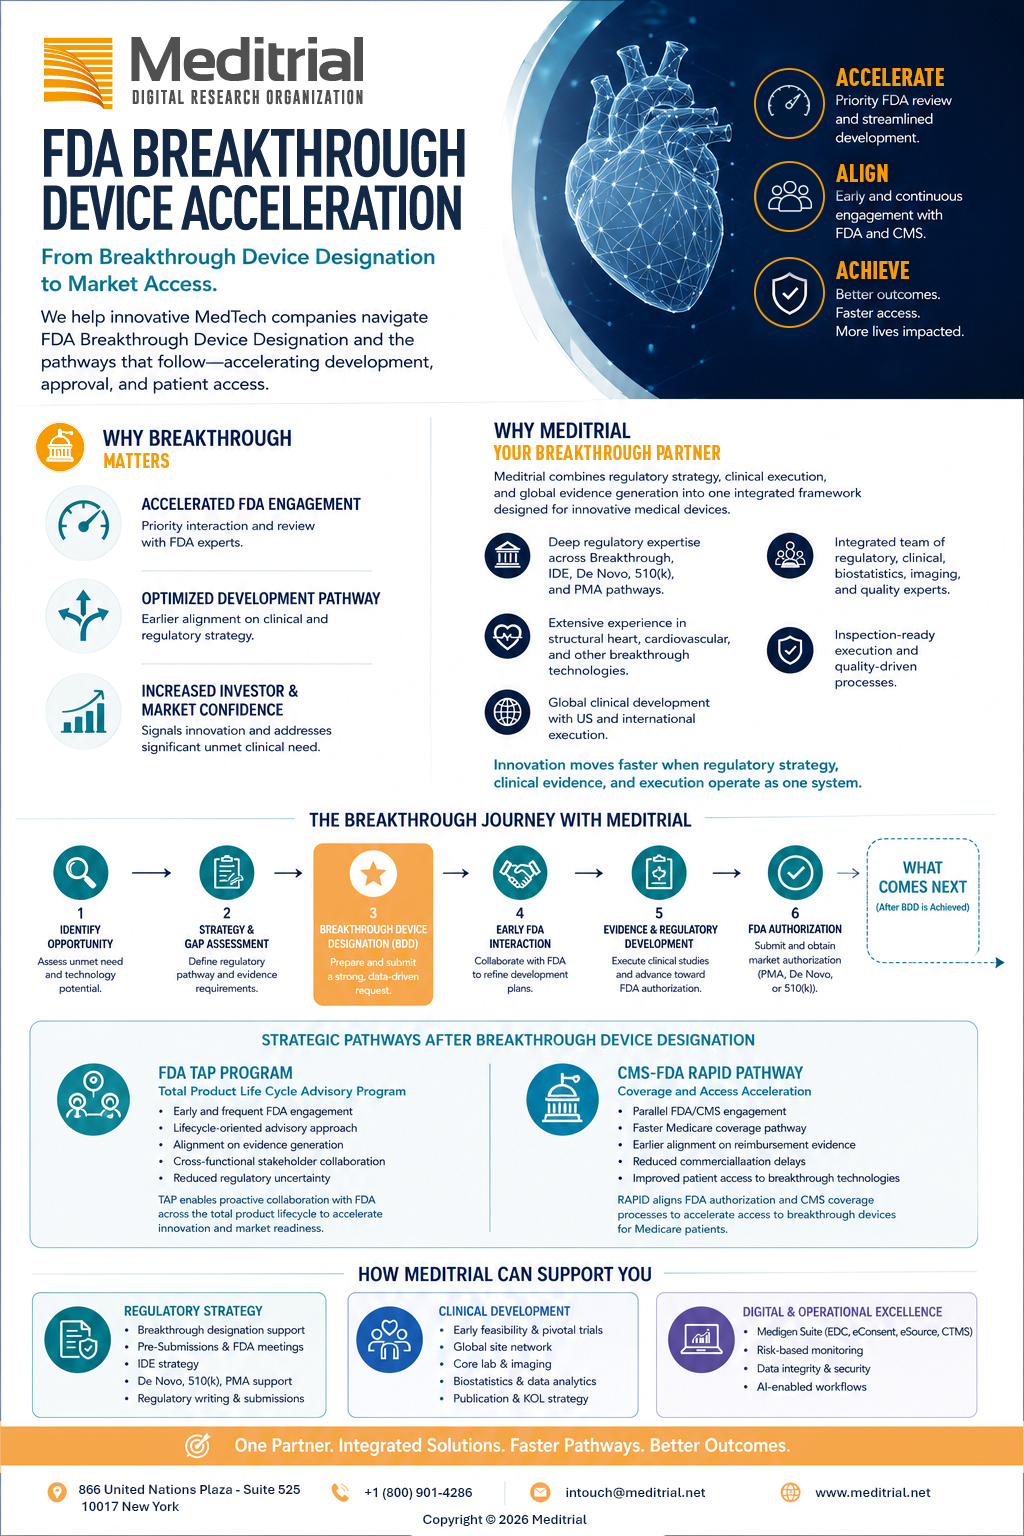
FDA Breakthrough Device Acceleration
How innovative MedTech companies can accelerate FDA approval, secure Medicare coverage, and reach patients faster—starting with Breakthrough Device Designation.
The contribution of patient associations in clinical trials
Historically, patient organizations have had the primary purpose of supporting the communities they represented. Today, this remains a crucial function, but modes have changed, as has the role that associations and patients themselves take on [...]
This article provides general information and does not constitute regulatory or legal advice. Requirements should be confirmed for the specific study, applicable regulatory framework, and markets concerned.