FDA issues guidance on Premarket Approval (PMA) and Humanitarian Device Exemption (HDE) supplements during COVID-19
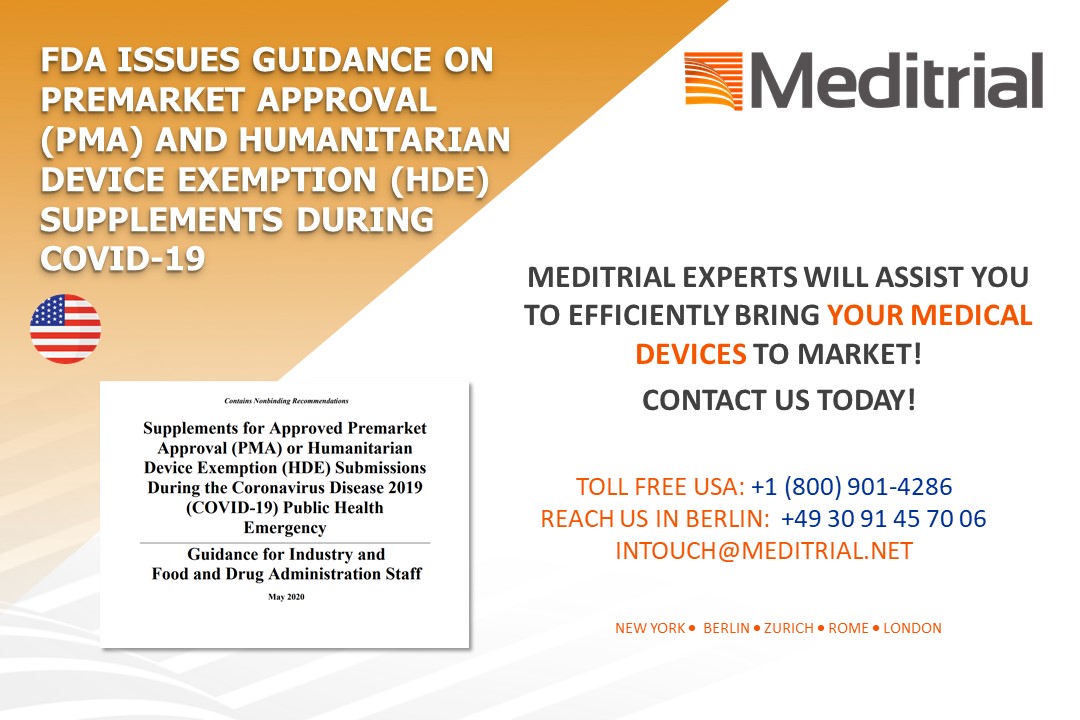

The US Food and Drug Administration (FDA) issued a guidance with immediate effect which allows medical device manufacturers to make “limited modifications” to their devices without submitting a premarket approval (PMA), humanitarian device exemption (HDE) supplement or a 30-day notice during the COVID-19 emergency.
The aim of this policy is to grant device manufacturers additional flexibility to address supply chain issues arising from material or component shortages or facility closures due to COVID-19-related disruptions.
“Due to these and other COVID-19 related developments, numerous manufacturers may need to make immediate changes such as adjusting manufacturing processes to allow for social distancing, adapting their manufacturing or design due to supply chain disruption, or to moving device production to a region that is less impacted by COVID-19.”
The policy does not apply to changes made to PMA or HDE devices that are unrelated to manufacturing limitations or supply chain issues caused by COVID-19 and do not extend to changes that have already been included in submissions to the agency.
The guidance includes a list of modifications that would typically require a supplement or notice that would generally be allowable under the policy, as well as a list of changes it says would create an undue risk and would not be allowable without prior submission of a PMA or HDE supplement or notice.
For more details, please see the guidance from FDA.
Meditrial can help – Contact us for immediate support
Meditrial has developed rapid workflows to conduct risk assessment, implement rescue plans and execute solutions for ongoing clinical trials:
- Identify patients requiring follow-up
- Conduct investigator and study nurse calls for training to address the situation
- Obtain authorizations for decentralized data collection
- Support to patients for alternative access to needed care
- Prepare procedures and worksheets for remote collection of patient clinical data
- Monitor work instructions and trainings for site support and remote monitoring
- EDC and ePRO solutions to enable study continuity
- Privacy protection measures
Contact Meditrial for immediate assistance in Europe or the US.
Table of Contents
Getting a CE mark for medical devices: a step-by-step guide
Practical guidance on the CE mark for medical devices under EU MDR, from classification and evidence to conformity assessment and EUDAMED
A Practical RBQM Framework for Medical Device Trials
Risk-based quality management for medical device trials, with practical actions for risk assessment, monitoring, data review, and trial oversight
EU Breakthrough Medical Devices Under MDCG 2025-9 (BtX)
EU breakthrough medical devices gain a clearer regulatory pathway under MDCG 2025-9, with early advice and lifecycle evidence planning
What Swissmedic Fast-Track Means for Clinical Trial Sponsors
Swissmedic swissethics Fast-Track Pilot Project accelerates eligible trial reviews and makes coordinated regulatory preparation increasingly important.
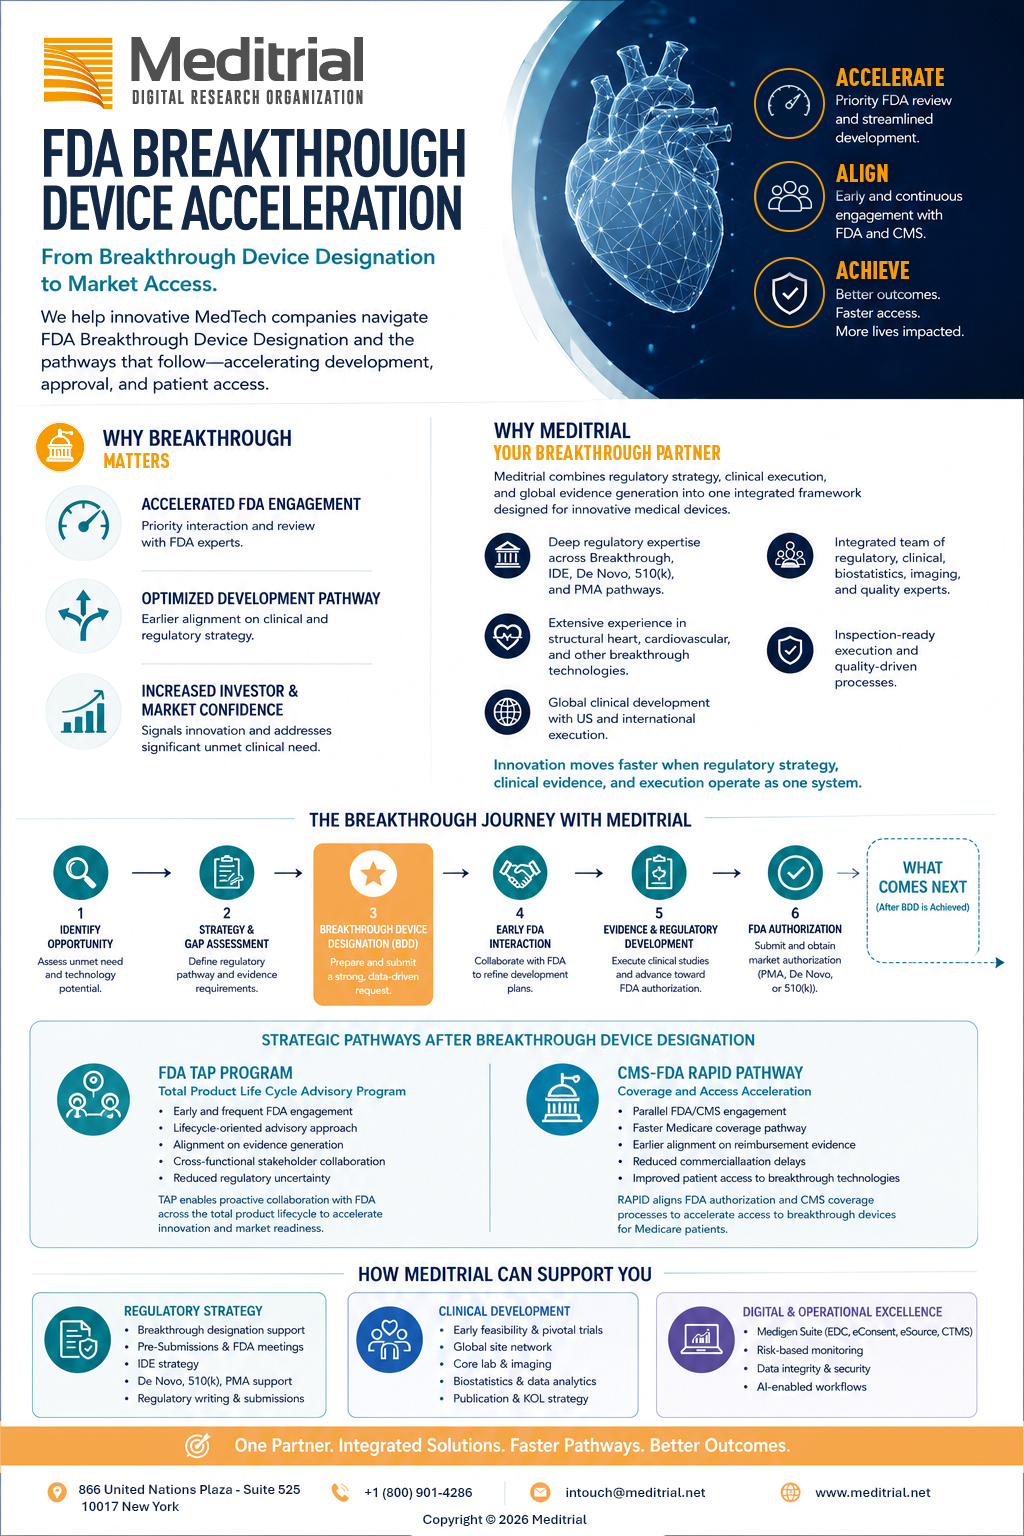
FDA Breakthrough Device Acceleration
How innovative MedTech companies can accelerate FDA approval, secure Medicare coverage, and reach patients faster—starting with Breakthrough Device Designation.
The contribution of patient associations in clinical trials
Historically, patient organizations have had the primary purpose of supporting the communities they represented. Today, this remains a crucial function, but modes have changed, as has the role that associations and patients themselves take on [...]
This article provides general information and does not constitute regulatory or legal advice. Requirements should be confirmed for the specific study, applicable regulatory framework, and markets concerned.