MDCG issued guidance on transitioning devices with ancillary substances to MDR
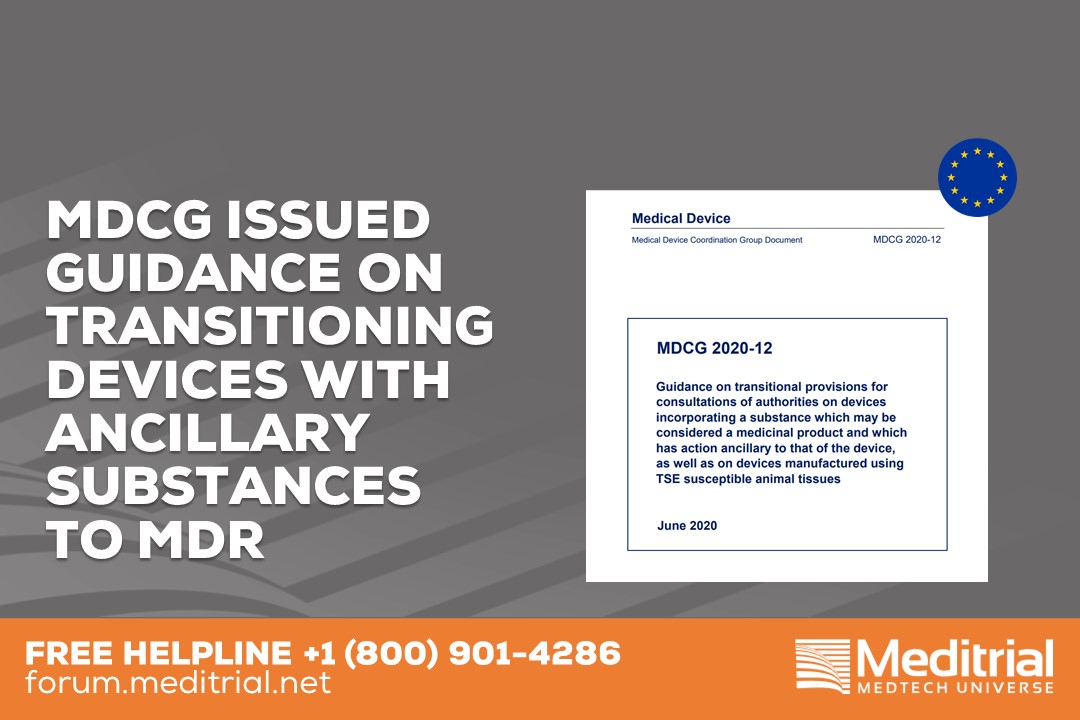

The European Commission’s Medical Device Coordination Group (MDCG) released guidance about the process for transitioning CE certificates for devices carrying ancillary substance which could be considered a medicinal product and those incorporating animal tissues to the MDR.
Ancillary substances
Medical devices that incorporate an ancillary substance that could be considered a medicinal product, must undergo a consultation with a medicinal product authority or the European Medicines Agency (EMA).
Following changes are introduced:
- a 210-day timeframe for the competent authority or EMA to provide an opinion for initial consultations and,
- a 60-day window for supplementary consultations
- notified bodies may not issue a certificate if the opinion is unfavorable.
Animal tissue
For devices manufactured using animal tissues, the first certification under the MDR requires the submission of a complete summary evaluation report to the competent authorities.
For more details, please see the guidance document from MDCG.
Meditrial has developed rapid workflows to conduct risk assessment, implement rescue plans and execute solutions for ongoing clinical trials:
- Identify patients requiring follow-up
- Conduct investigator and study nurse calls for training to address the situation
- Obtain authorizations for decentralized data collection
- Support to patients for alternative access to needed care
- Prepare procedures and worksheets for remote collection of patient clinical data
- Monitor work instructions and trainings for site support and remote monitoring
- EDC and ePRO solutions to enable study continuity
- Privacy protection measures
Contact Meditrial for immediate assistance in Europe or the US.
Table of Contents
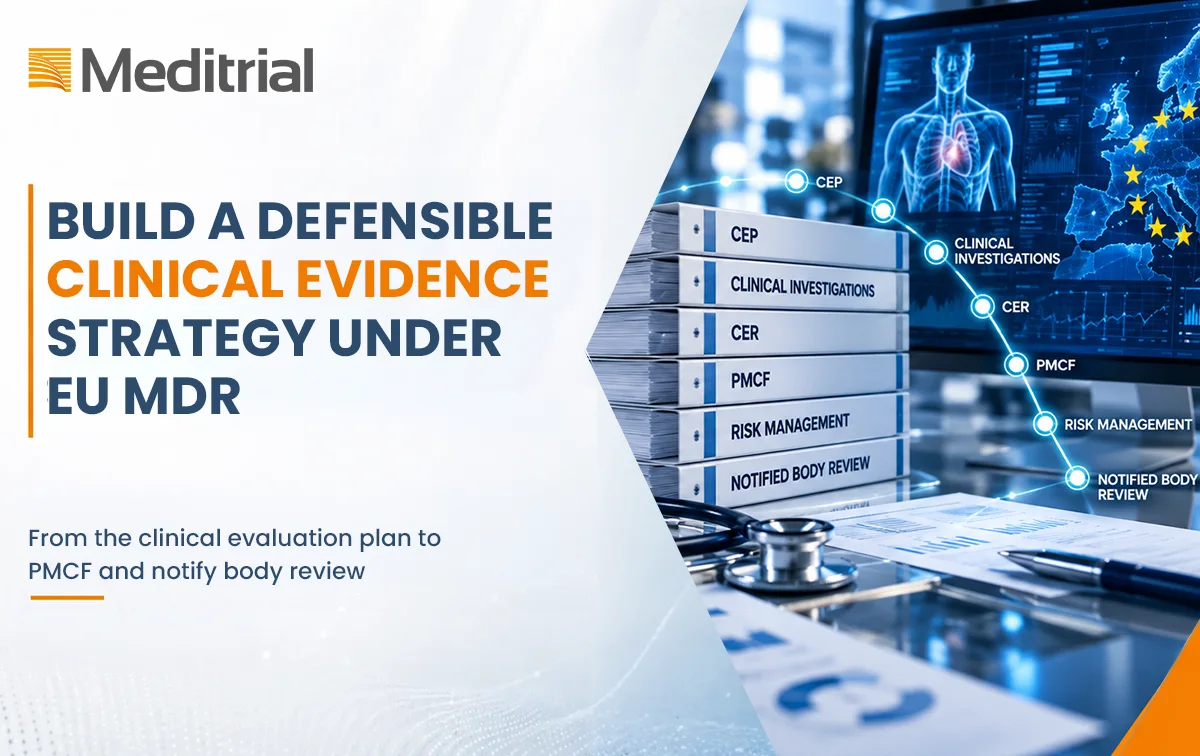
Build a Defensible Clinical Evidence Strategy Under EU MDR
Build a defensible clinical evidence strategy under MDR Annex XIV, from the clinical evaluation plan to PMCF and notified body review.
Getting a CE mark for medical devices: a step-by-step guide
Practical guidance on the CE mark for medical devices under EU MDR, from classification and evidence to conformity assessment and EUDAMED
A Practical RBQM Framework for Medical Device Trials
Risk-based quality management for medical device trials, with practical actions for risk assessment, monitoring, data review, and trial oversight
EU Breakthrough Medical Devices Under MDCG 2025-9 (BtX)
EU breakthrough medical devices gain a clearer regulatory pathway under MDCG 2025-9, with early advice and lifecycle evidence planning
What Swissmedic Fast-Track Means for Clinical Trial Sponsors
Swissmedic swissethics Fast-Track Pilot Project accelerates eligible trial reviews and makes coordinated regulatory preparation increasingly important.
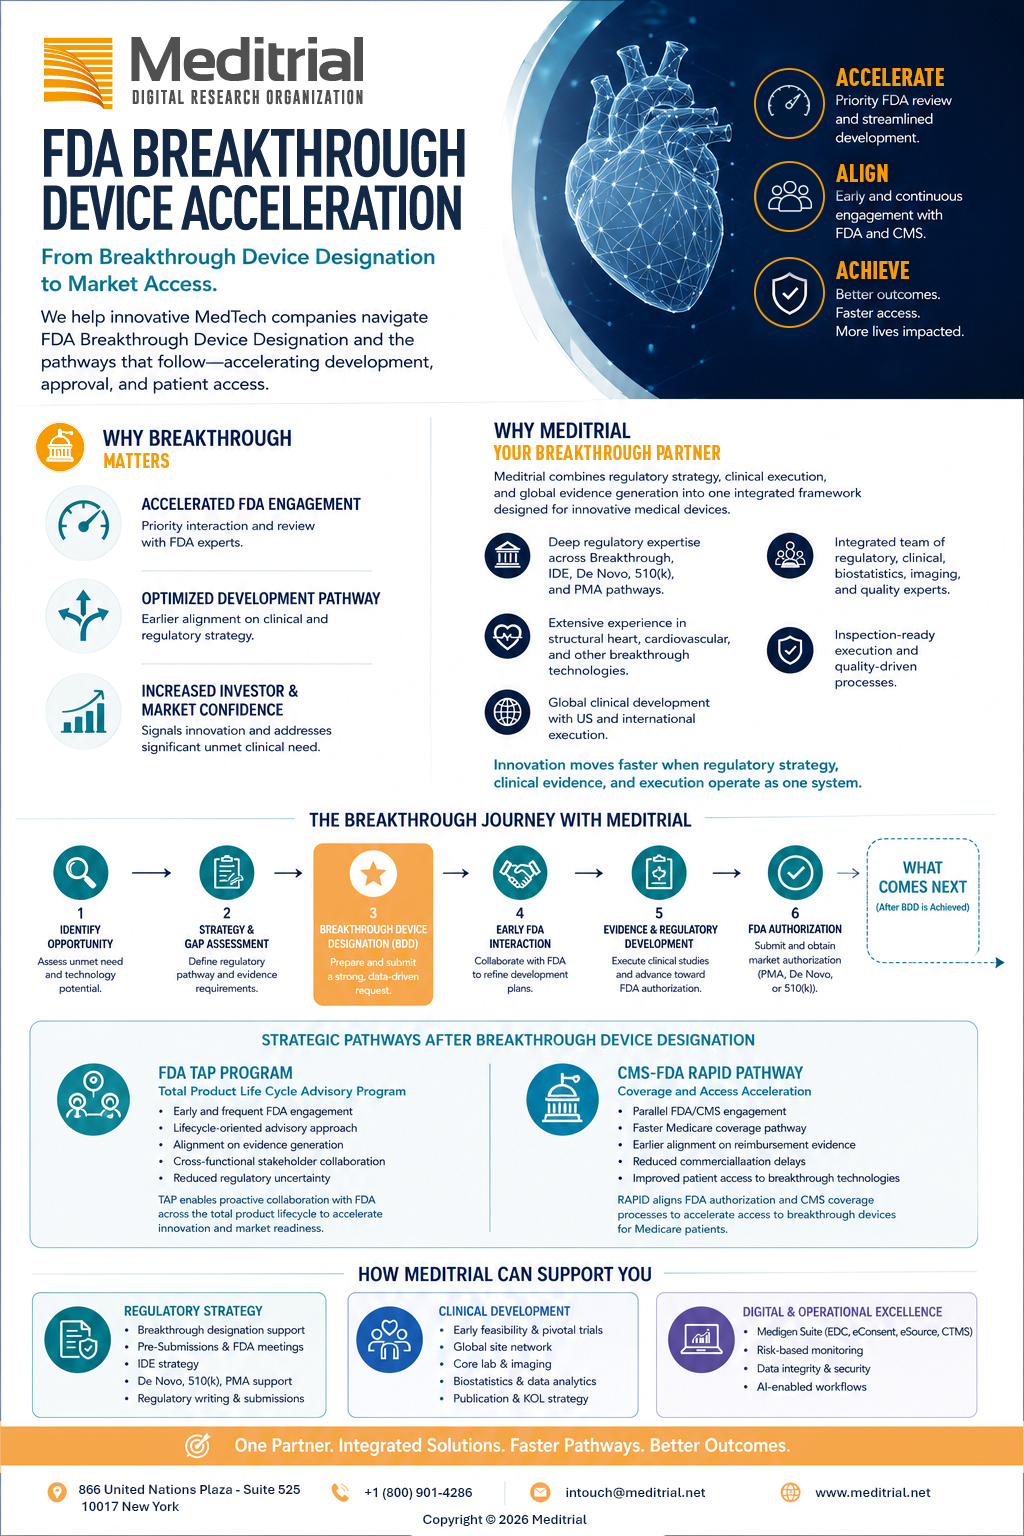
FDA Breakthrough Device Acceleration
How innovative MedTech companies can accelerate FDA approval, secure Medicare coverage, and reach patients faster—starting with Breakthrough Device Designation.
This article provides general information and does not constitute regulatory or legal advice. Requirements should be confirmed for the specific study, applicable regulatory framework, and markets concerned.