The USA: FDA is holding back the enforcement of certain UDI requirements

The US Food and Drug Administration (FDA) has published final guidance for unique device identification compliance dates for Class I and Unclassified Medical Devices. This immediately in effect guidance also clarifies the Agency policy concerning compliance dates for certain Devices requiring direct marking.
The compliance dates established for class I and unclassified devices, other than implantable, life-supporting, or life-sustaining (I/LS/LS) devices are:
- September 24, 2018, for the following requirements:
- Standard date formatting (21 CFR 801.18),
- Labeling (21 CFR 801.20, 21 CFR 801.50), and
- Global Unique Device Identification Database (GUDID) data submission (21 CFR 830.300); and
- September 24, 2020, for direct mark requirements (21 CFR 801.45).
“This guidance also describes FDA’s direct mark compliance policy for class III, LS/LS, and class II devices that are non-sterile, that are manufactured and labeled prior to their applicable direct mark compliance date, and that remain in inventory, as well as for class I and unclassified devices that are not LS/LS devices, that are non-sterile, that are manufactured and labeled prior to September 24, 2022, and that remain in inventory.”
FDA recognized that “preparing to implement UDI requirements while addressing the challenges related to Coronavirus Disease 2019 (COVID-19) could be very difficult and could divert resources from COVID-19 response efforts.” Therefore, it indicates “To the extent this policy helps labelers remain focused on public health needs related to COVID-19, we believe the policy is further consistent with the public health.”
For more information, please see the guidance document from the FDA.
Contact Meditrial for immediate assistance in the US or in Europe.
Table of Contents
Getting a CE mark for medical devices: a step-by-step guide
Practical guidance on the CE mark for medical devices under EU MDR, from classification and evidence to conformity assessment and EUDAMED
A Practical RBQM Framework for Medical Device Trials
Risk-based quality management for medical device trials, with practical actions for risk assessment, monitoring, data review, and trial oversight
EU Breakthrough Medical Devices Under MDCG 2025-9 (BtX)
EU breakthrough medical devices gain a clearer regulatory pathway under MDCG 2025-9, with early advice and lifecycle evidence planning
What Swissmedic Fast-Track Means for Clinical Trial Sponsors
Swissmedic swissethics Fast-Track Pilot Project accelerates eligible trial reviews and makes coordinated regulatory preparation increasingly important.
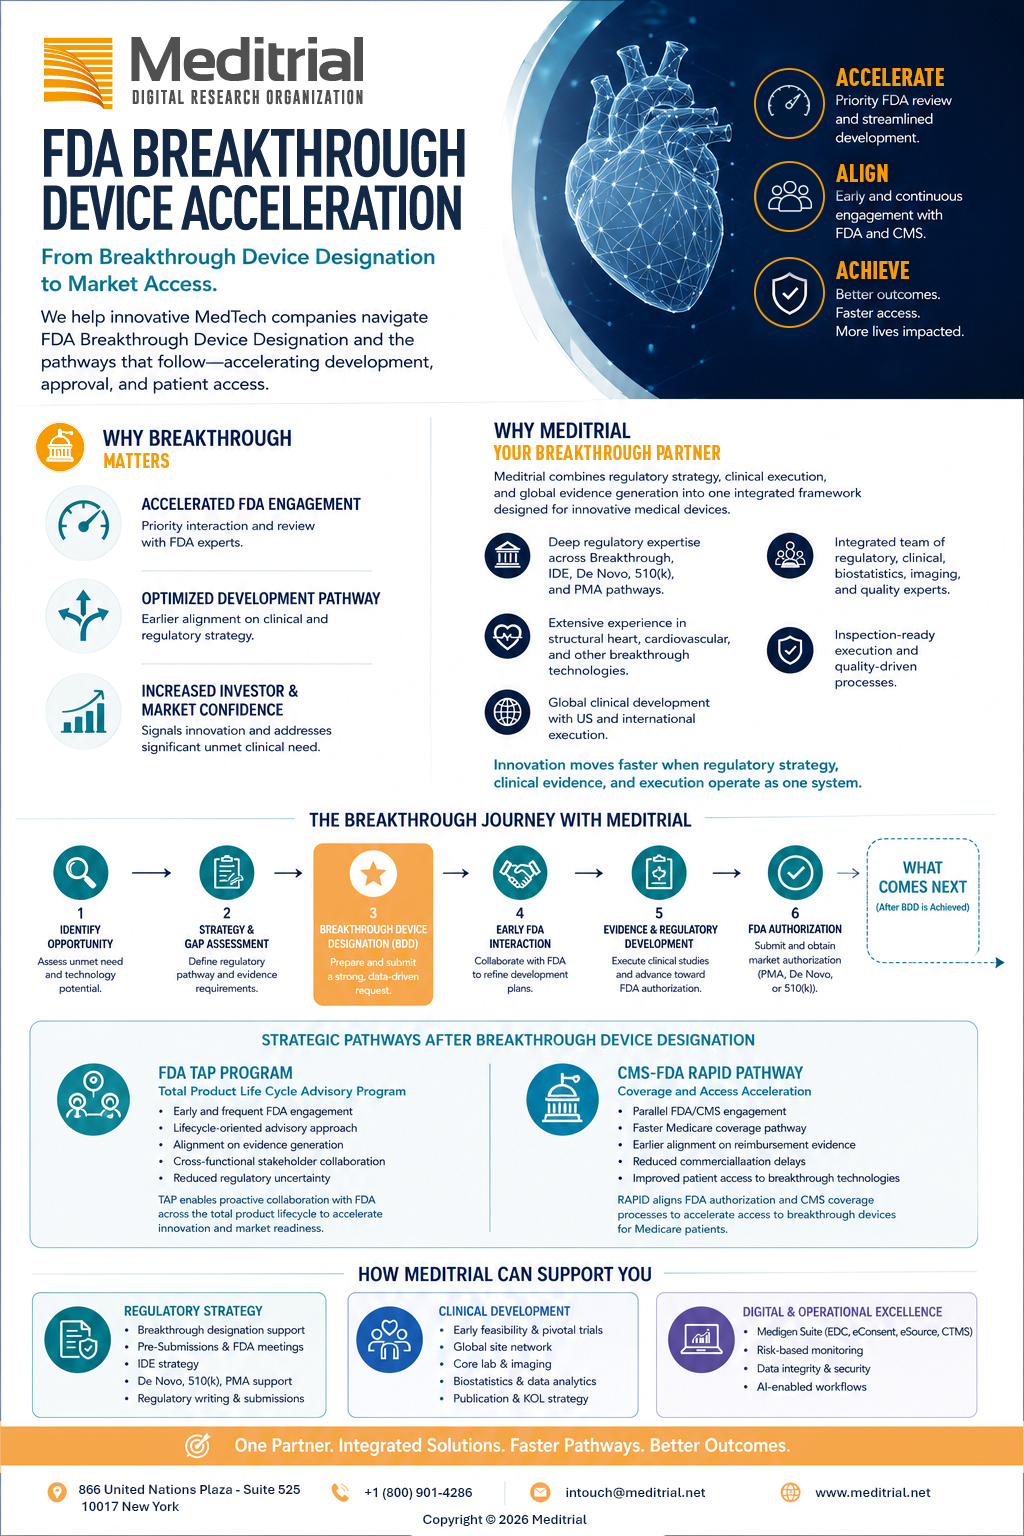
FDA Breakthrough Device Acceleration
How innovative MedTech companies can accelerate FDA approval, secure Medicare coverage, and reach patients faster—starting with Breakthrough Device Designation.
The contribution of patient associations in clinical trials
Historically, patient organizations have had the primary purpose of supporting the communities they represented. Today, this remains a crucial function, but modes have changed, as has the role that associations and patients themselves take on [...]
This article provides general information and does not constitute regulatory or legal advice. Requirements should be confirmed for the specific study, applicable regulatory framework, and markets concerned.