US: FDA “green lights” the path for ePRO through regulation
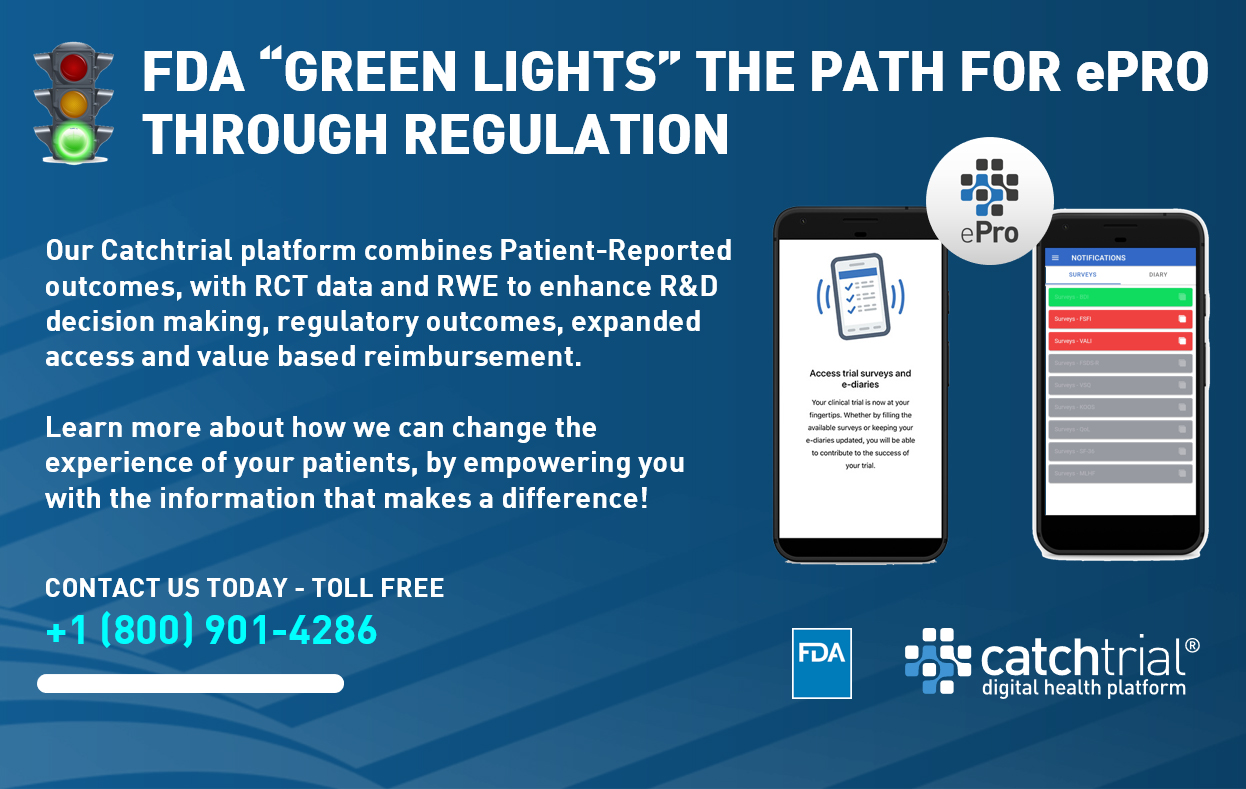

The FDA issued a guidance for the selection of patient-reported outcome (PRO) instruments in clinical evaluation of their products. The guidance details such topics as modification or adaptation of existing PRO instruments as well as the possibility of including patient-reported outcome instruments within real-world data sources.
As explained by the FDA in the draft guidance, PRO instruments “facilitate the systematic collection of how patients feel, function and survive as valid scientific evidence to support the regulatory and healthcare decision-making process.”
Catchtrial ePRO App
OUR Catchtrial ePRO App facilitates better interaction between the patient, investigator and sponsor, ensures faster trial completion, and significant cost savings. It works with any desktop or mobile device, including Android and Apple iOS. ePRO simply allows you to stay tuned with patient health.
ePRO App features
• Manage images transfer in the virtual trial thanks to smartphone camera control
• GDPR and HIPAA compliant – no patient PHI entry required, no personal data breach
• Off-line data entry in case of no internet connection
Catchtrial electronic Patient-Reported Outcomes (ePRO) is a survey of the status of a patient’s health that comes directly from the patient, i.e. the patient reports the data directly.
These surveys can include psychological symptoms, patient diary and general quality of life measures, such as how the condition impacts a patient’s daily life.
Learn more about Catchtrial ePRO App!
For more details, please see the guidance from the FDA.
Contact Meditrial for immediate assistance in Europe or the US.
Table of Contents
Practical 510(k) vs PMA vs De Novo FDA Pathway Guide
Practical 510(k) vs PMA vs De Novo guidance for choosing an FDA device pathway based on classification, predicates, risk, and evidence needs
Build a Defensible Clinical Evidence Strategy Under EU MDR
Build a defensible clinical evidence strategy under MDR Annex XIV, from the clinical evaluation plan to PMCF and notified body review.
Getting a CE mark for medical devices: a step-by-step guide
Practical guidance on the CE mark for medical devices under EU MDR, from classification and evidence to conformity assessment and EUDAMED
A Practical RBQM Framework for Medical Device Trials
Risk-based quality management for medical device trials, with practical actions for risk assessment, monitoring, data review, and trial oversight
EU Breakthrough Medical Devices Under MDCG 2025-9 (BtX)
EU breakthrough medical devices gain a clearer regulatory pathway under MDCG 2025-9, with early advice and lifecycle evidence planning
What Swissmedic Fast-Track Means for Clinical Trial Sponsors
Swissmedic swissethics Fast-Track Pilot Project accelerates eligible trial reviews and makes coordinated regulatory preparation increasingly important.
This article provides general information and does not constitute regulatory or legal advice. Requirements should be confirmed for the specific study, applicable regulatory framework, and markets concerned.