EU: EMA expands the use of remote source data verification in clinical trials
The European Commission and European Medicines Agency’s updated the guidance on managing clinical trials conducted during the COVID-19 pandemic by expanding the use and scope of remote source data verification (rSDV) in clinical trials.
While previous versions of the guidance stated that rSDV would “only be considered necessary for very few trials” involving the treatment or prevention of COVID-19, “Or in the final data cleaning steps before database lock in pivotal trials investigating serious or life-threatening conditions with no satisfactory treatment option.”, the updated guidance has expanded the types of trials that may use rSDV.
According to the guidance, rSDV can be used for trials:
- involving the treatment or prevention of COVID-19
- investigating serious or life-threatening conditions
- pivotal trials
- “where the absence of SDV for critical data may likely pose unacceptable risks to participants’ safety or the reliability/integrity of trial results”
- “involving particularly vulnerable participants such as children or those temporarily […] or permanently […] incapable of giving their informed consent.”
Moreover, the updated Annex 1 clarifies that the principal investigator (PI), the PI’s institution and the sponsor may be jointly responsible as controllers for ensuring that participants’ information is safeguarded.
For more details, please see the official guidance from the EMA and the European Commission.
- Identify patients requiring follow-up
- Conduct investigator and study nurse calls for training to address the situation
- Obtain authorizations for decentralized data collection
- Support to patients for alternative access to needed care
- Prepare procedures and worksheets for remote collection of patient clinical data
- Monitor work instructions and trainings for site support and remote monitoring
- EDC and ePRO solutions to enable study continuity
- Privacy protection measures
Contact Meditrial for immediate assistance in Europe or the US.
Table of Contents
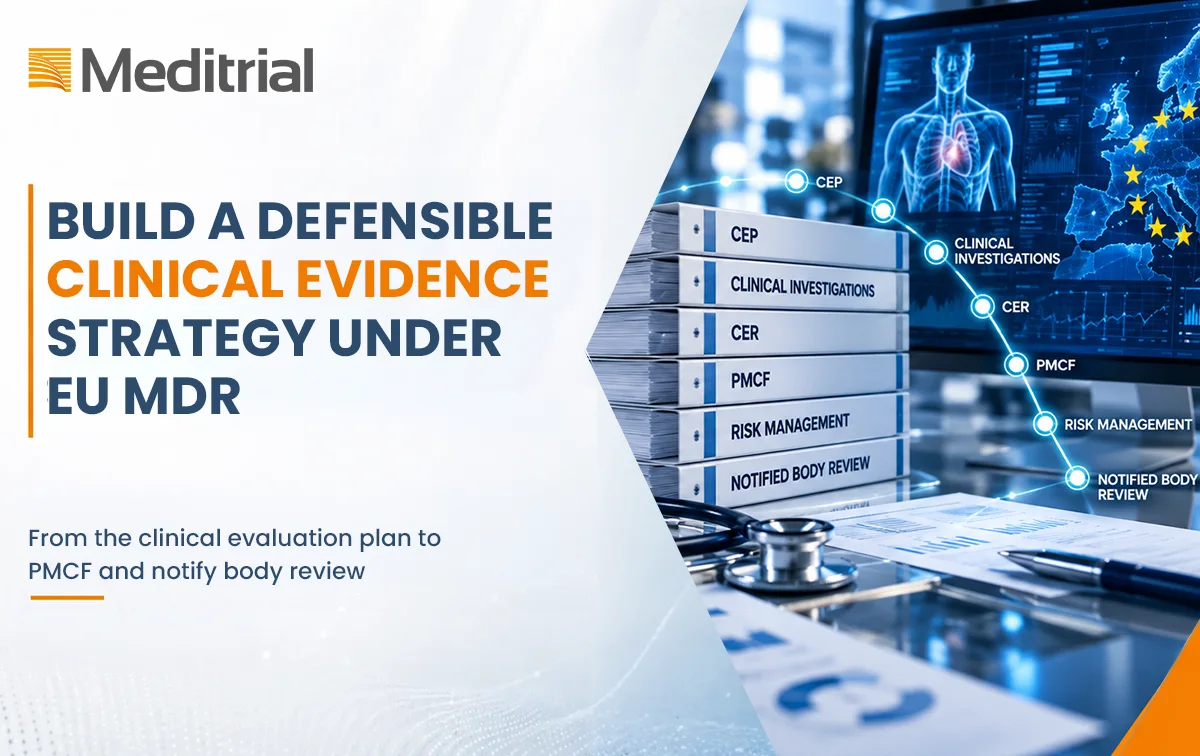
Build a Defensible Clinical Evidence Strategy Under EU MDR
Build a defensible clinical evidence strategy under MDR Annex XIV, from the clinical evaluation plan to PMCF and notified body review.
Getting a CE mark for medical devices: a step-by-step guide
Practical guidance on the CE mark for medical devices under EU MDR, from classification and evidence to conformity assessment and EUDAMED
A Practical RBQM Framework for Medical Device Trials
Risk-based quality management for medical device trials, with practical actions for risk assessment, monitoring, data review, and trial oversight
EU Breakthrough Medical Devices Under MDCG 2025-9 (BtX)
EU breakthrough medical devices gain a clearer regulatory pathway under MDCG 2025-9, with early advice and lifecycle evidence planning
What Swissmedic Fast-Track Means for Clinical Trial Sponsors
Swissmedic swissethics Fast-Track Pilot Project accelerates eligible trial reviews and makes coordinated regulatory preparation increasingly important.
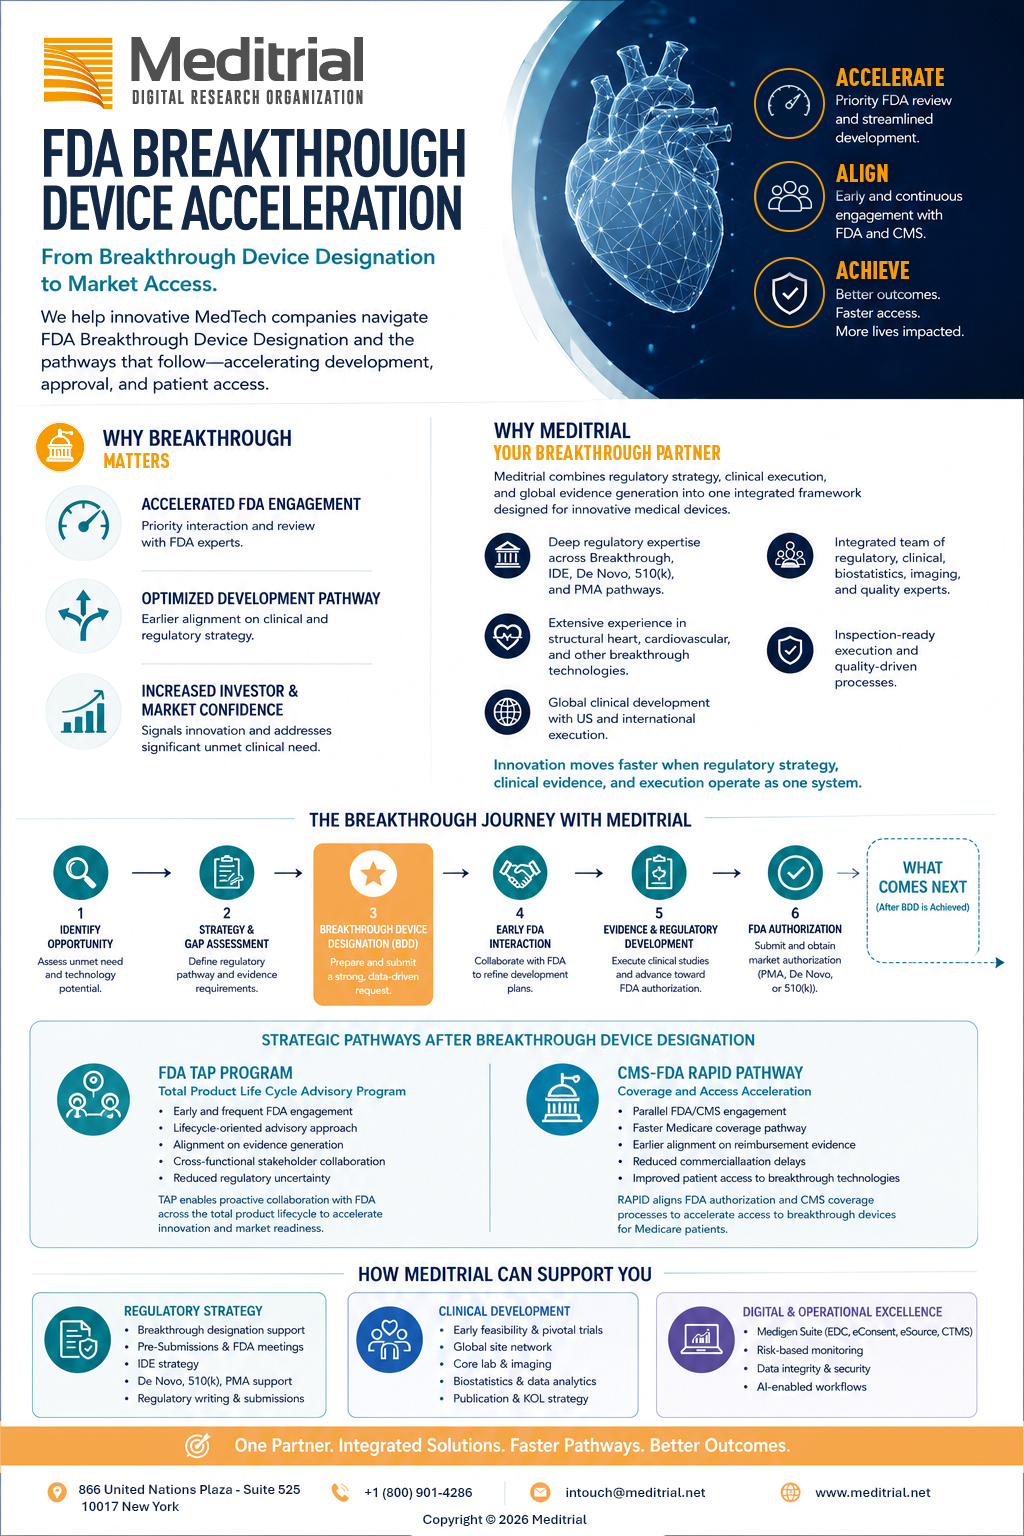
FDA Breakthrough Device Acceleration
How innovative MedTech companies can accelerate FDA approval, secure Medicare coverage, and reach patients faster—starting with Breakthrough Device Designation.
This article provides general information and does not constitute regulatory or legal advice. Requirements should be confirmed for the specific study, applicable regulatory framework, and markets concerned.