EU: The European Commission issues a guidance on management of legacy devices in Eudamed
The European Commission issued a new guidance explaining how legacy devices and in vitro diagnostics (IVDs) will be managed in its Eudamed database, expected to be fully rolled out by May 2022.
“Legacy Devices are defined as Medical Devices, Active Implantable Medical Devices and In Vitro Diagnostic Medical Devices that are covered by a valid certificate issued in accordance with Directive 93/42/EEC, Directive 90/385/EEC or Directive 98/79/EC and that continue to be placed on the market after the date of application of Regulation (EU) 2017/745 (MDR) or Regulation 2017/746 (IVDR).”
In general, the Commission says that, “Legacy Devices are subject to the MDR or IVRD registration requirements with some exceptions such as the assignment of a Basic [universal device identification-device identifier] UDI-DI and a UDI-DI.”
The document also explains how to link “regulation devices” to legacy devices after they become compliant with MDR or IVDR and provides information about the format of the Eudamed DI when it is not generated from a UDI-DI.
Do you know how to register for EUDAMED?
Contact Meditrial to learn everything you need to know on how to prepare! With us, you will be the first to comply.
Read Meditrial’s whitepaper to learn more on the MDR and Eudamed.
For more details, please see the guidance from the European Commission.
Contact Meditrial for immediate assistance in Europe or the US.
Table of Contents
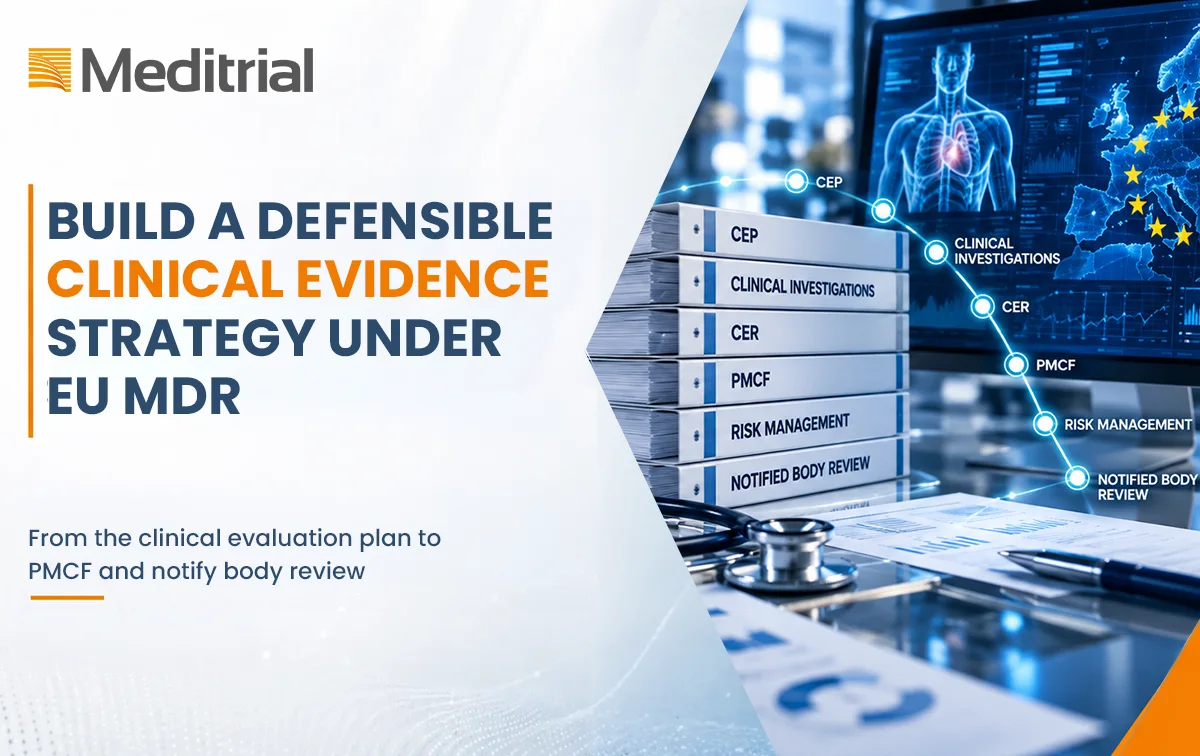
Build a Defensible Clinical Evidence Strategy Under EU MDR
Build a defensible clinical evidence strategy under MDR Annex XIV, from the clinical evaluation plan to PMCF and notified body review.
Getting a CE mark for medical devices: a step-by-step guide
Practical guidance on the CE mark for medical devices under EU MDR, from classification and evidence to conformity assessment and EUDAMED
A Practical RBQM Framework for Medical Device Trials
Risk-based quality management for medical device trials, with practical actions for risk assessment, monitoring, data review, and trial oversight
EU Breakthrough Medical Devices Under MDCG 2025-9 (BtX)
EU breakthrough medical devices gain a clearer regulatory pathway under MDCG 2025-9, with early advice and lifecycle evidence planning
What Swissmedic Fast-Track Means for Clinical Trial Sponsors
Swissmedic swissethics Fast-Track Pilot Project accelerates eligible trial reviews and makes coordinated regulatory preparation increasingly important.
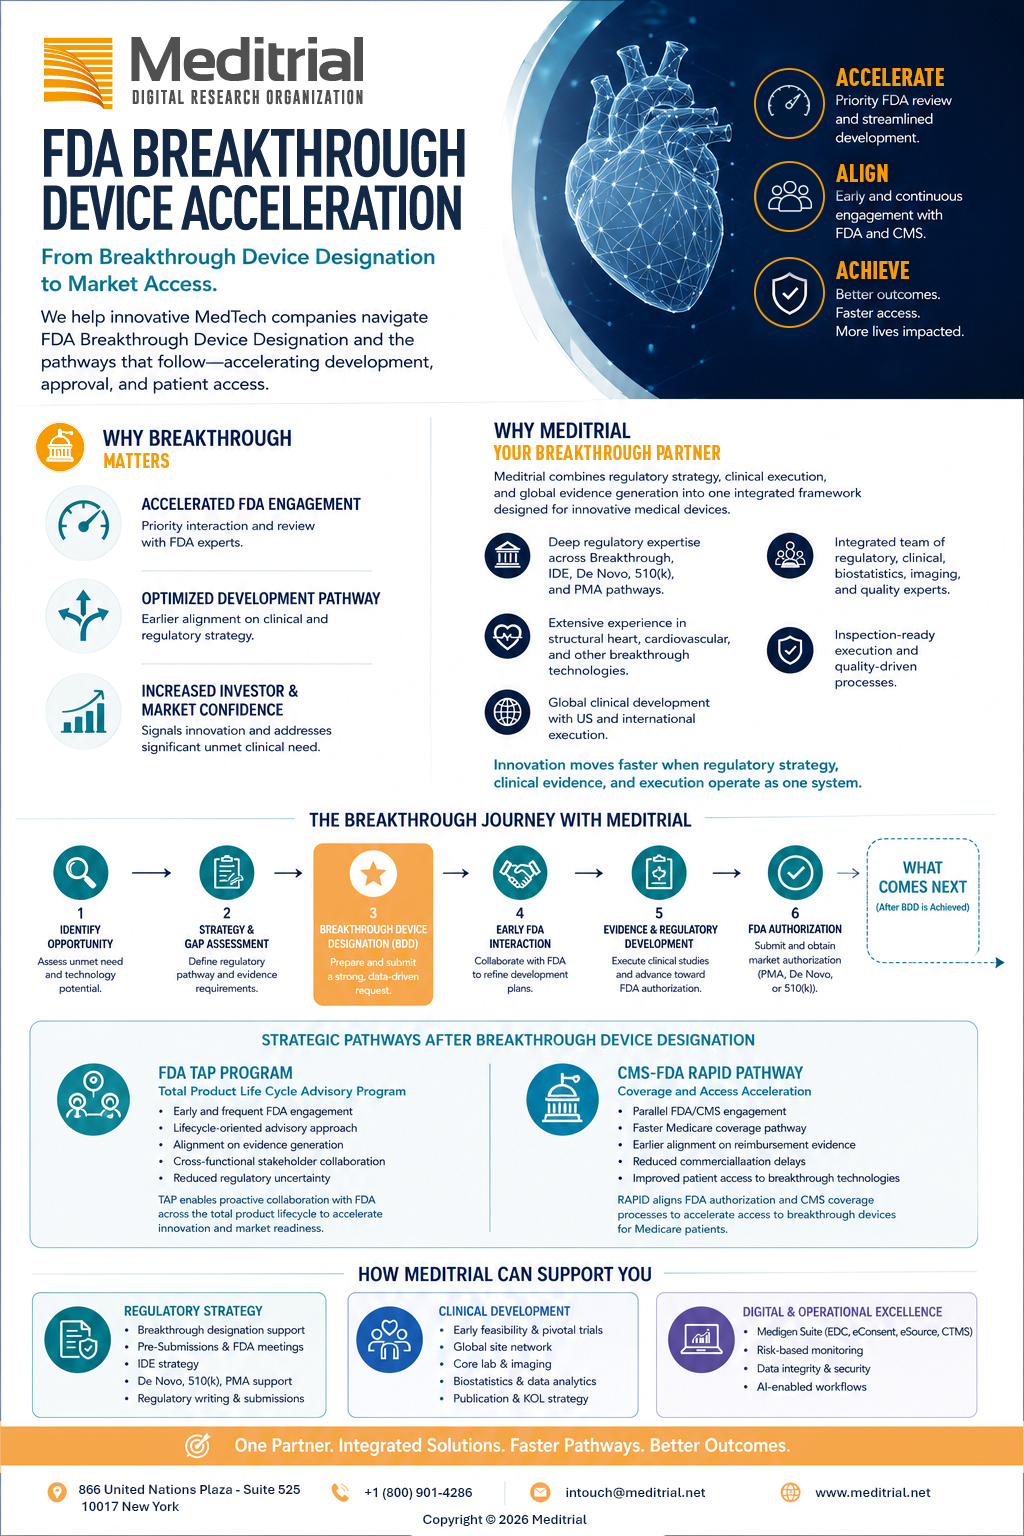
FDA Breakthrough Device Acceleration
How innovative MedTech companies can accelerate FDA approval, secure Medicare coverage, and reach patients faster—starting with Breakthrough Device Designation.
This article provides general information and does not constitute regulatory or legal advice. Requirements should be confirmed for the specific study, applicable regulatory framework, and markets concerned.