EU: The MDCG informs on how to handle class D IVDs during transition to new regulation IVDR
The transition period to the new in vitro diagnostic regulation (IVDR) in the EU will last till next 26 May, 2022. In order to inform device manufacturers, the European Commission´s Medical Device Coordination Group issued a Q&A including information on how to handle class D devices, the highest risk class, during the transition period.
Contact Meditrial for a free consultation regarding your IVD! With international presence and complete range of services, Meditrial is a leading Clinical Research Organization (CRO), a full-service partner for clinical investigations, digital clinical solutions and market access pathways.
“According to Regulation (EU) 2017/746 on in vitro diagnostic medical devices (the IVDR), as part of conformity assessment of class D in vitro diagnostic medical devices (IVDs), the manufacturer must submit an application to a notified body. In addition to the assessment by the notified body, under certain conditions particular elements may be reviewed by an expert panel and/or tested by an EU reference laboratory (EURL). The establishment of EURLs for a range of class D devices has been identified as an important priority by the Medical Device Coordination Group. The European Commission is preparing the implementing acts on tasks and criteria of the EURLs and fees they may levy, referred to in Article 100(8) of the IVDR, as well as the first call for applications to be submitted by the Member States. This document provides indications for how to apply the IVDR provisions related to expert panels and EURLs during the transition period, i.e. before 26 May 2022.”
MDCG 2021-4 Application of transitional provisions for certification of class D in vitro diagnostic medical devices according to Regulation (EU) 2017/746, April 2021
For more details, please see the document from the MDCG.
Contact Meditrial for immediate assistance in Europe or the US.
Table of Contents
Getting a CE mark for medical devices: a step-by-step guide
Practical guidance on the CE mark for medical devices under EU MDR, from classification and evidence to conformity assessment and EUDAMED
A Practical RBQM Framework for Medical Device Trials
Risk-based quality management for medical device trials, with practical actions for risk assessment, monitoring, data review, and trial oversight
EU Breakthrough Medical Devices Under MDCG 2025-9 (BtX)
EU breakthrough medical devices gain a clearer regulatory pathway under MDCG 2025-9, with early advice and lifecycle evidence planning
What Swissmedic Fast-Track Means for Clinical Trial Sponsors
Swissmedic swissethics Fast-Track Pilot Project accelerates eligible trial reviews and makes coordinated regulatory preparation increasingly important.
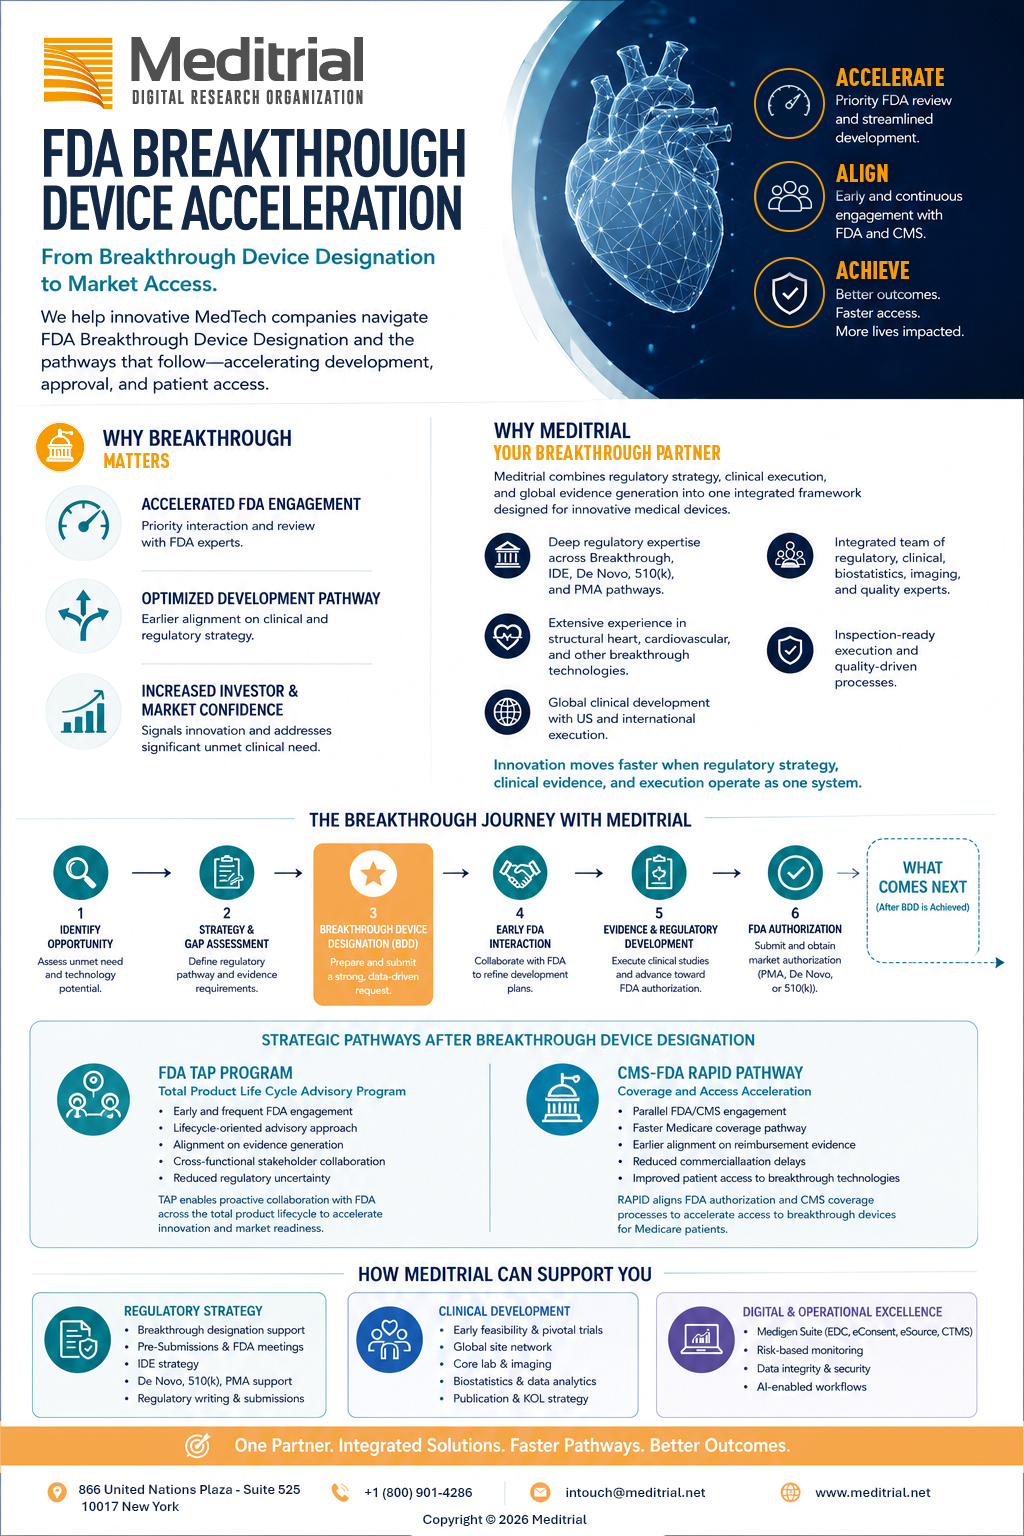
FDA Breakthrough Device Acceleration
How innovative MedTech companies can accelerate FDA approval, secure Medicare coverage, and reach patients faster—starting with Breakthrough Device Designation.
The contribution of patient associations in clinical trials
Historically, patient organizations have had the primary purpose of supporting the communities they represented. Today, this remains a crucial function, but modes have changed, as has the role that associations and patients themselves take on [...]
This article provides general information and does not constitute regulatory or legal advice. Requirements should be confirmed for the specific study, applicable regulatory framework, and markets concerned.