MDCG outlines rules on EUDAMED registration for custom made and legacy devices
The European Commission’s Medical Device Coordination Group (MDCG) last week published a new guidance with Q&A on obligations and related rules for the registration in EUDAMED of actors not subject to registration requirements under the Art. 31 Medical Device Regulation (MDR) and Art. 28 In Vitro Diagnostic Regulation (IVDR). These rules apply also to manufacturers of custom-made and legacy devices.
Contact Meditrial to receive help with EUDAMED registration! We will help you comply with MDR and IVDR.
Under new Regulations, actors other than manufacturers, authorized representatives and importers are required to register as actors in EUDAMED. Registration procedures are expected to be laid out in an upcoming Commission Implementing Regulation.
🔸 Difference between issuing an Actor ID and SRN number
Actor ID is applied to all actors in EUDAMED for their identification and is generated automatically when the registration is approved by the relevant competent authority. In contrast, the SRN is assigned in EUDAMED specifically to manufacturers, authorized representatives and importers by a competent authority.
🔸 Manufacturers of only custom-made devices
Manufacturers who only produce custom-made devices are exempted from registry requirements before placing their devices on the market. However, to enter information on class III custom-made implantable devices and to report serious incidents, field safety corrective actions, filed safety notices and any trends of non-serious incidents, manufacturers of custom-made devices will need to register as actors in EUDAMED. Also all non-EU manufacturers of only custom-made devices and their authorized representatives are required to register.
🔸 Manufacturers and authorized representatives of only legacy devices
Manufacturers of only legacy devices must register as actors in EUDAMED within six months of the date of publication of the notice under Article 34(3) of MDR announcing the functionality of EUDAMED. This requirement affects non-EU manufacturers and their authorized representatives, who will be assigned an Actor ID that is not an SRN.
🔸 Manufacturers of only “old” devices, which are made available on the Union market and/or are still in use
Manufacturers of “old” devices, which include devices placed on the market prior to, or according to, the medical device and in vitro diagnostic directives, will need to register in EUDAMED to submit reports of serious incidents or FSCAs. This duty also applies to non-EU manufacturers and authorized representatives. However, inactive manufacturers and authorized representatives of old devices are not subject to EUDAMED registration requirements.
🔸 System and procedure pack producers (SPPP)
The system and procedure pack producers (SPPPs) must register as an actor within six months of the notice announcing the functionality of EUDAMED, as they are obligated to assign a basic unique device identification-device identifier (UDI-DI) to be provided to the UDI database.
For more details, please see the MDCG Guidance.
Receive dedicated support! Contact Meditrial for immediate assistance.
Table of Contents
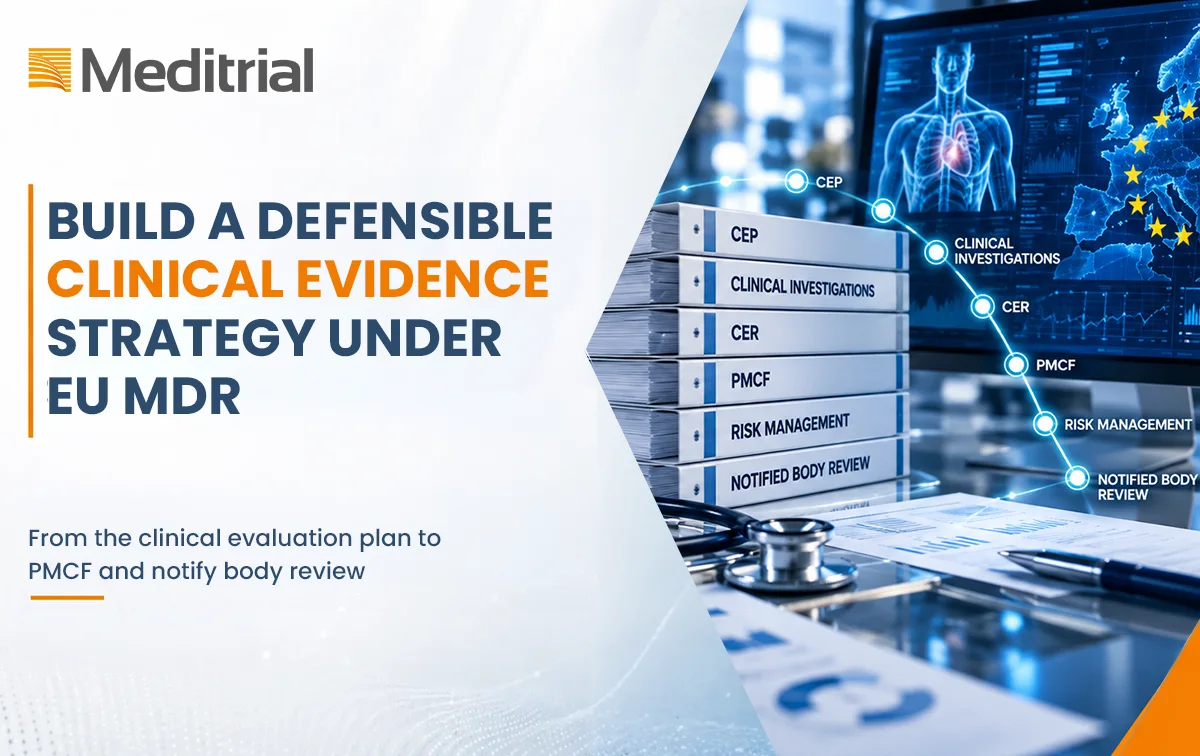
Build a Defensible Clinical Evidence Strategy Under EU MDR
Build a defensible clinical evidence strategy under MDR Annex XIV, from the clinical evaluation plan to PMCF and notified body review.
Getting a CE mark for medical devices: a step-by-step guide
Practical guidance on the CE mark for medical devices under EU MDR, from classification and evidence to conformity assessment and EUDAMED
A Practical RBQM Framework for Medical Device Trials
Risk-based quality management for medical device trials, with practical actions for risk assessment, monitoring, data review, and trial oversight
EU Breakthrough Medical Devices Under MDCG 2025-9 (BtX)
EU breakthrough medical devices gain a clearer regulatory pathway under MDCG 2025-9, with early advice and lifecycle evidence planning
What Swissmedic Fast-Track Means for Clinical Trial Sponsors
Swissmedic swissethics Fast-Track Pilot Project accelerates eligible trial reviews and makes coordinated regulatory preparation increasingly important.
FDA Breakthrough Device Acceleration
How innovative MedTech companies can accelerate FDA approval, secure Medicare coverage, and reach patients faster—starting with Breakthrough Device Designation.
This article provides general information and does not constitute regulatory or legal advice. Requirements should be confirmed for the specific study, applicable regulatory framework, and markets concerned.