New MDCG guidance 2022-4
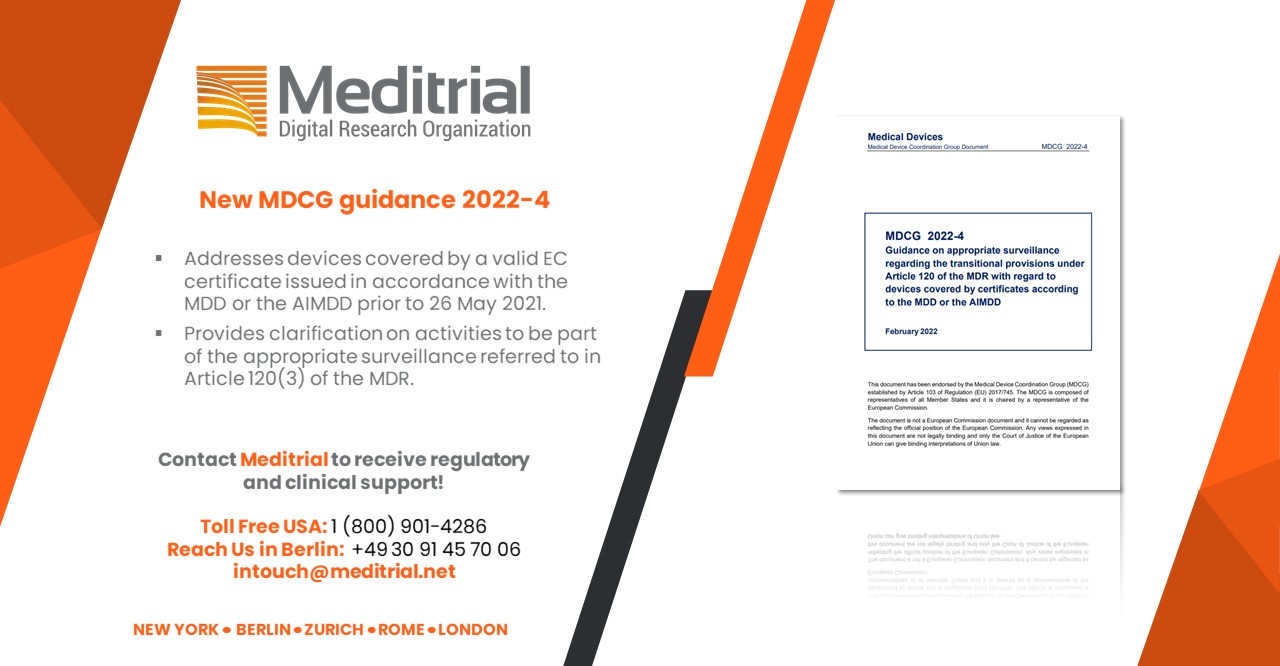
The new MDCG guidance 2022-4 has been published. It clarifies how Notified Bodies shall perform surveillance of legacy devices during the transition period defined in MDR Article 120.
The document provides guidance for surveillance activities according to four different scenarios:
- Manufacturers of legacy devices that have not applied for certification under the MDR,
- Manufacturers of legacy devices and MDR devices having already implemented the MDR requirements in their systems and whose application for MDR certification is being reviewed by the Notified Body having issued the MDD/AIMDD certificate,
- Manufacturers of legacy devices and MDR devices already certified by the same Notified Body under MDR,
- Manufacturers of legacy devices and MDR devices already certified by another Notified Body under the MDR.
In addition, for class IIa and IIb MDD devices, the technical documentation assessments will continue on a sampling basis according to the existing sampling plan generated by the Notified Body under the MDD.
Annex of the document includes a comparison table – quality management system requirements in the #MDD and the MDR.
Article 110(3) of Regulation (EU) 2017/746 (IVDR) is outside the scope of this guidance document. However, principles outlined in the text and that are common between EU legislations for medical devices and in-vitro diagnostic medical devices may be applied also for activities to be performed by notified bodies according to Article 110(3) of the IVDR.
👉🏻 Read more in the MDCG Guidance.
Receive regulatory and clinical support from Meditrial experts. Get in touch today!
Table of Contents
Build a Defensible Clinical Evidence Strategy Under EU MDR
Build a defensible clinical evidence strategy under MDR Annex XIV, from the clinical evaluation plan to PMCF and notified body review.
Getting a CE mark for medical devices: a step-by-step guide
Practical guidance on the CE mark for medical devices under EU MDR, from classification and evidence to conformity assessment and EUDAMED
A Practical RBQM Framework for Medical Device Trials
Risk-based quality management for medical device trials, with practical actions for risk assessment, monitoring, data review, and trial oversight
EU Breakthrough Medical Devices Under MDCG 2025-9 (BtX)
EU breakthrough medical devices gain a clearer regulatory pathway under MDCG 2025-9, with early advice and lifecycle evidence planning
What Swissmedic Fast-Track Means for Clinical Trial Sponsors
Swissmedic swissethics Fast-Track Pilot Project accelerates eligible trial reviews and makes coordinated regulatory preparation increasingly important.
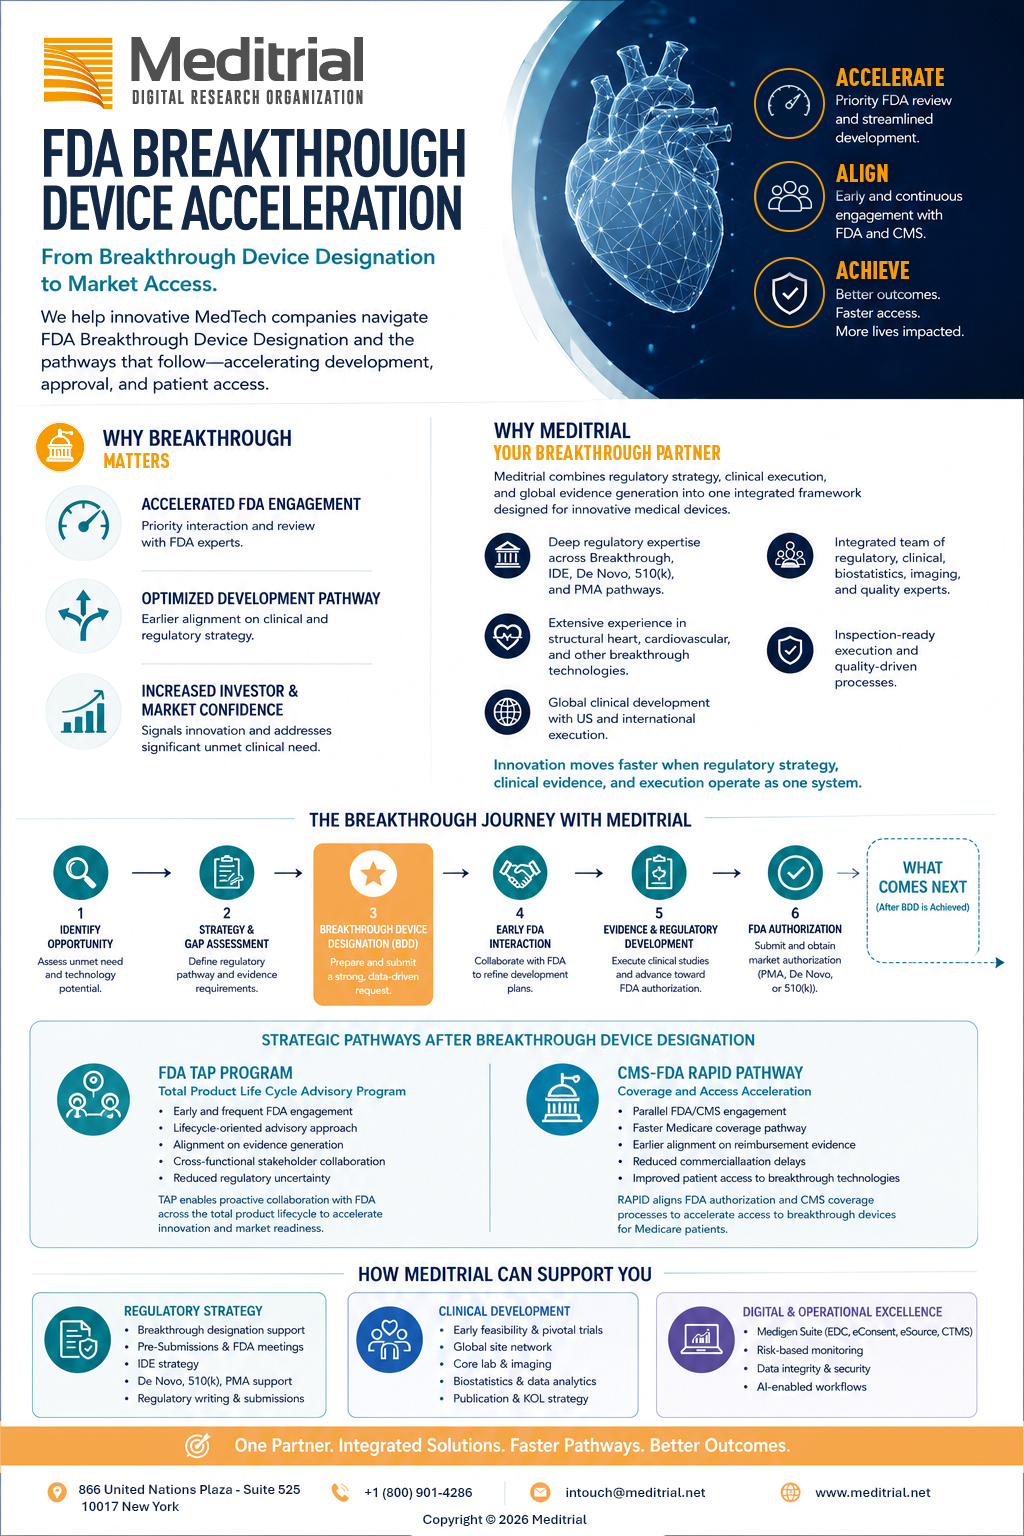
FDA Breakthrough Device Acceleration
How innovative MedTech companies can accelerate FDA approval, secure Medicare coverage, and reach patients faster—starting with Breakthrough Device Designation.
This article provides general information and does not constitute regulatory or legal advice. Requirements should be confirmed for the specific study, applicable regulatory framework, and markets concerned.