FDA Finalizes Guidance for Premarket Pathways for Combination Products

The FDA has finalized guidance on the use of premarket pathways for combination products.
The document defines combination products, the coordination between the FDA and sponsors, and recommendations on how to determine the types of premarket submissions for a particular product. Annex of the document contains specific examples of how the guidance may be applied.
Meditrial team of medical, regulatory and technical experts will assist you to efficiently bring innovative combination products to market.
What are combination products?
The guidance defines combination products as consisting of two or more different types of drugs, devices, or biological products. These components may be contained in a single unit, such as a drug-eluting stent packaged as a first-aid kit with an antiseptic drug, or they could be individual products that are intended for use together and are cross-labeled, such as a light-emitting device with a light-activated drug.
Determining center jurisdiction
The FDA intends to assign each combination product to an FDA center that will have primary jurisdiction. The lead center will be based on which component provides the mode of action “expected to make the greatest contribution to the overall intended therapeutic effects” of the combination product, known as the primary mode of action (PMOA).
Types of marketing applications
Premarket review of a combination product generally occurs under a single application. The marketing application type submitted should generally coincide with the PMOA of the combination product (a PMA, De Novo, or 510(k) for a device-led combination product; an NDA or ANDA for a drug-led combination product; or a BLA for a biologic-led combination product).
Although single applications are appropriate in most cases, the FDA guidance emphasizes that sponsors should provide data on all components of a combination product. “To appropriately ensure the safety and effectiveness of a combination product in a single application, such application should enable a substantially similar evaluation to that which would be applied to each constituent part if they were reviewed under separate applications (e.g., an ANDA or NDA for a drug and a PMA, De Novo, or 510(k) for a device), including consideration of data and information that would be reviewed under the separate applications.”
Read more: https://www.fda.gov/media/119958/download
Meditrial accelerates the pathway to approval for life-changing devices, therapies and health technologies. Contact us today!
Table of Contents
Build a Defensible Clinical Evidence Strategy Under EU MDR
Build a defensible clinical evidence strategy under MDR Annex XIV, from the clinical evaluation plan to PMCF and notified body review.
Getting a CE mark for medical devices: a step-by-step guide
Practical guidance on the CE mark for medical devices under EU MDR, from classification and evidence to conformity assessment and EUDAMED
A Practical RBQM Framework for Medical Device Trials
Risk-based quality management for medical device trials, with practical actions for risk assessment, monitoring, data review, and trial oversight
EU Breakthrough Medical Devices Under MDCG 2025-9 (BtX)
EU breakthrough medical devices gain a clearer regulatory pathway under MDCG 2025-9, with early advice and lifecycle evidence planning
What Swissmedic Fast-Track Means for Clinical Trial Sponsors
Swissmedic swissethics Fast-Track Pilot Project accelerates eligible trial reviews and makes coordinated regulatory preparation increasingly important.
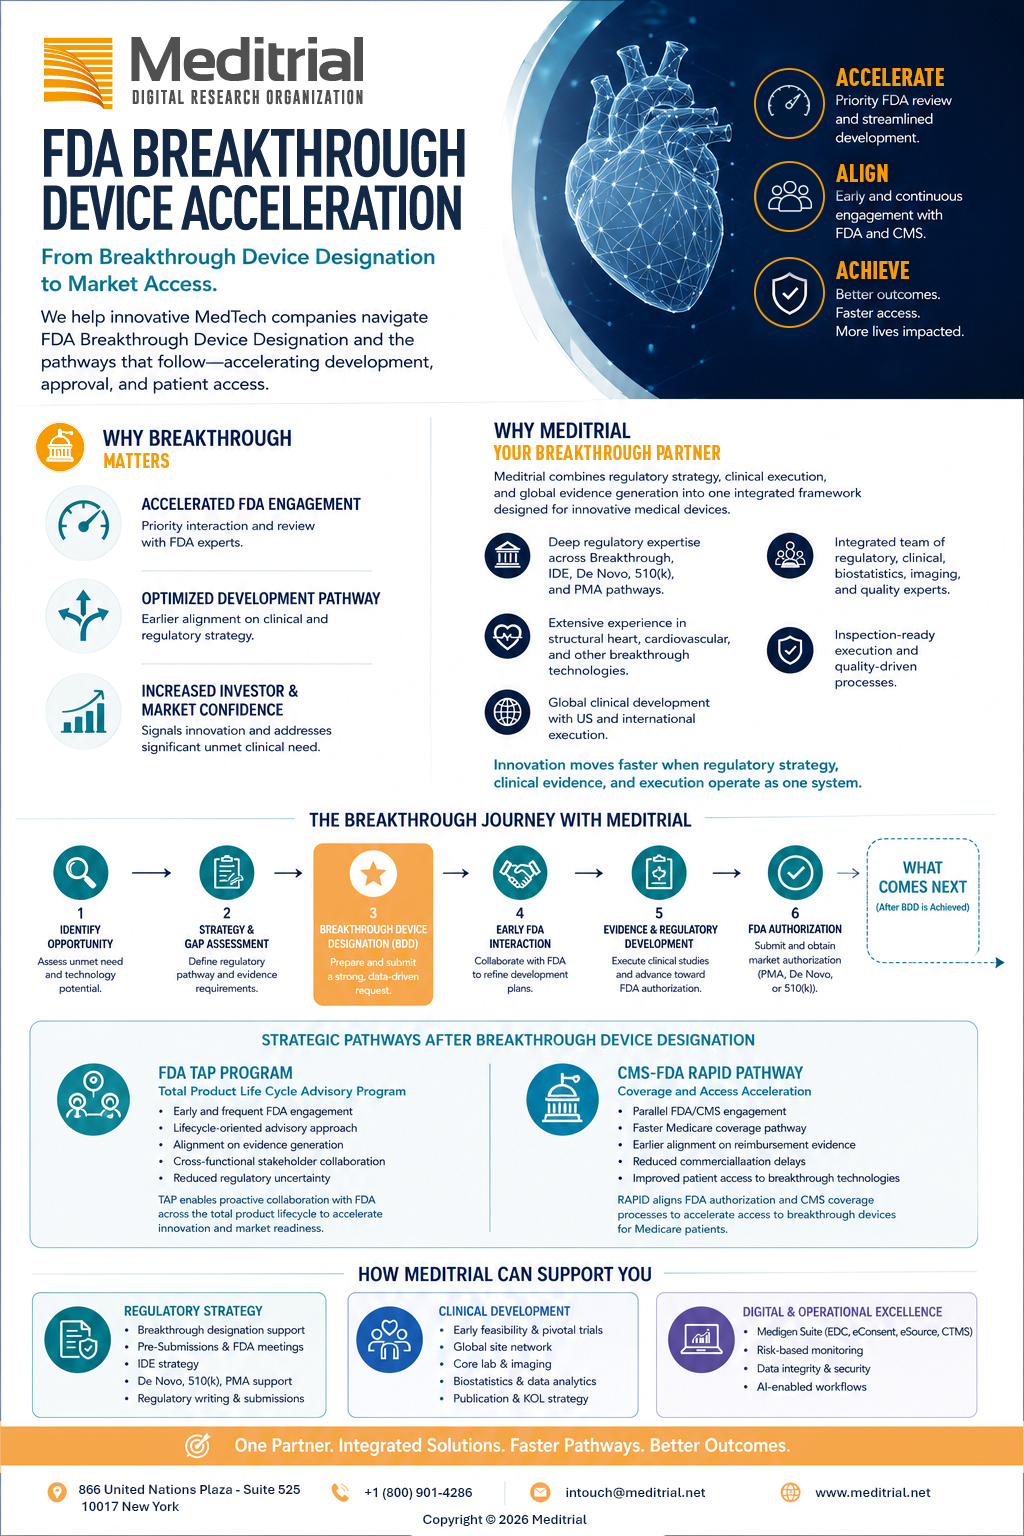
FDA Breakthrough Device Acceleration
How innovative MedTech companies can accelerate FDA approval, secure Medicare coverage, and reach patients faster—starting with Breakthrough Device Designation.
This article provides general information and does not constitute regulatory or legal advice. Requirements should be confirmed for the specific study, applicable regulatory framework, and markets concerned.