UPGRADE YOUR TRIALS FOR MDR BY MAY 26, 2021!
WHY WE
MUST UPDATE
OUR TRIALS
If you sponsor a trial that is already ongoing, you must update your safety procedures in accordance with the MDR by the deadline of May 26, 2021.
If you are just in the process of submitting a new trial, but it has not started yet, the Authorities will expect that you submit it in compliance with the MDR.
What does it mean to you?
- Your ongoing studies will need an amendment to gain approval for the changes
- Your planning and timelines will be affected by the new application procedures
- Your safety management must consider new definitions, process and timelines
- You will need to upgrade the essential study documents (protocol, IB, informed consent) to comply with new requirements.
Are you ready?
As the implementation of the EU MDR has already proven to be a complex and challenging process, it is important to ensure you are going in the right direction. For example, there are increased requirements surrounding the study design justification, greater focus on risk management, and more emphasis on patient rights and privacy.
Contact us now
- Partnering with Meditrial will allow you a quick and rapid upgrade of your trials to the MDR.
- All of your questions will be answered by regulatory experts in simplified language
- You will have access to dedicated tutorials and educational materials
- Our medical writers will provide assistance with the document revisions
- Our clinical team will help to implement the revised study.
Meditrial will not only ensure your medical device complies at every stage of its life cycle, but that you ultimately achieve market approval.
To learn more, visit: www.meditrial.net
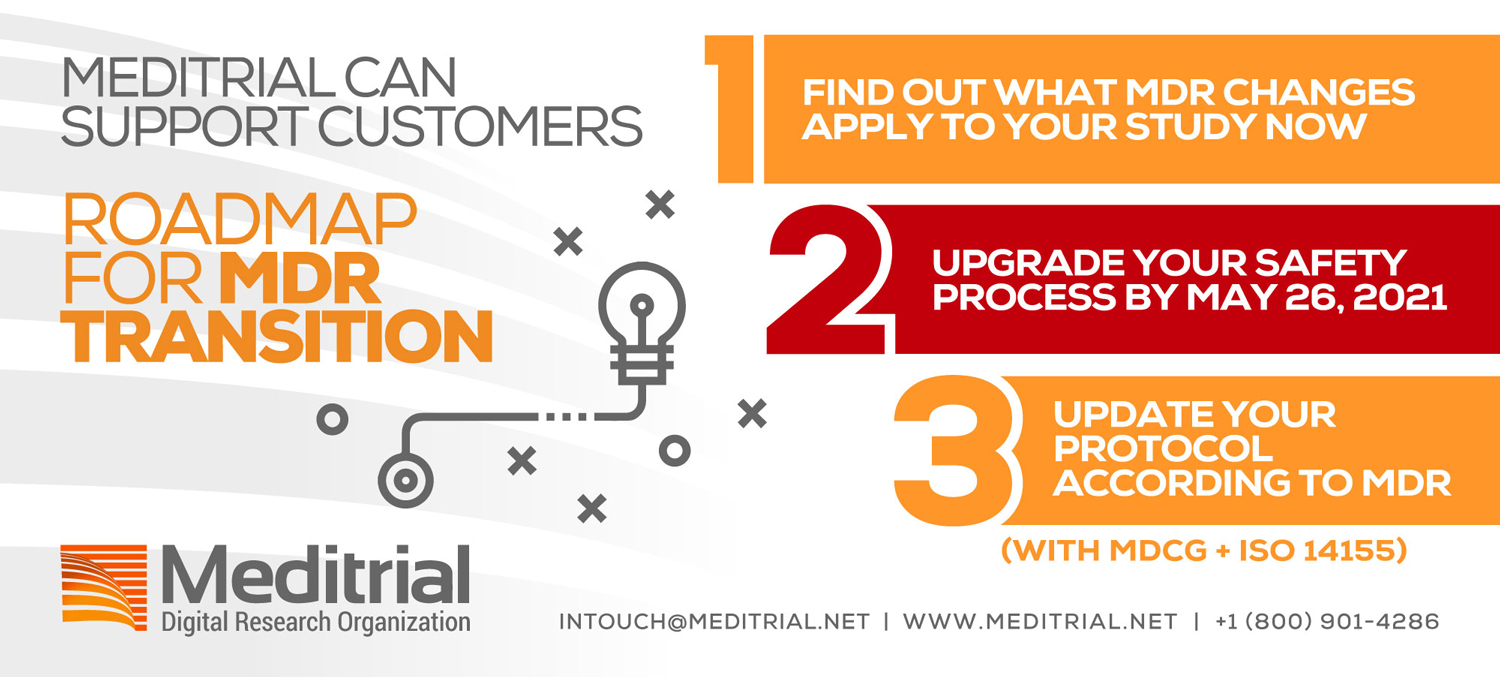
Table of Contents
Practical 510(k) vs PMA vs De Novo FDA Pathway Guide
Practical 510(k) vs PMA vs De Novo guidance for choosing an FDA device pathway based on classification, predicates, risk, and evidence needs
Build a Defensible Clinical Evidence Strategy Under EU MDR
Build a defensible clinical evidence strategy under MDR Annex XIV, from the clinical evaluation plan to PMCF and notified body review.
Getting a CE mark for medical devices: a step-by-step guide
Practical guidance on the CE mark for medical devices under EU MDR, from classification and evidence to conformity assessment and EUDAMED
A Practical RBQM Framework for Medical Device Trials
Risk-based quality management for medical device trials, with practical actions for risk assessment, monitoring, data review, and trial oversight
EU Breakthrough Medical Devices Under MDCG 2025-9 (BtX)
EU breakthrough medical devices gain a clearer regulatory pathway under MDCG 2025-9, with early advice and lifecycle evidence planning
What Swissmedic Fast-Track Means for Clinical Trial Sponsors
Swissmedic swissethics Fast-Track Pilot Project accelerates eligible trial reviews and makes coordinated regulatory preparation increasingly important.
This article provides general information and does not constitute regulatory or legal advice. Requirements should be confirmed for the specific study, applicable regulatory framework, and markets concerned.