FDA-EMA Parallel Scientific Advice Pilot Program for Complex Generic/Hybrid Products
The US Food and Drug Administration (FDA) and the European Medicines Agency (EMA) launched a new pilot program to provide parallel scientific advice (PSA) to sponsors of complex generic drugs, or hybrid medicines, as they are called in the EU.
“The goal of the PSA program is to provide a mechanism for EMA and FDA to concurrently consider and jointly exchange with applicants the agencies’ views on scientific questions during the development phase of hybrid/complex generic products.” Moreover, the aim of the program is to increase dialogue between the two agencies and applicants from the beginning of the product development.
Meditrial team of medical, regulatory and technical experts will assist you to efficiently bring innovative medicines to market. Please feel free to contact us for advice and support.
Some examples of studies that may benefit from parallel scientific advice in the complex generic/hybrid product space include “comparative non-clinical and comparative clinical studies involving innovative bioequivalence study designs and the use of methodologies such as modeling and simulation.”
PSA program is voluntary and will only consider products that meet both the FDA’s definition of complex generic and EMA’s definition of a hybrid medicine as the regulatory definitions differ between the two jurisdictions.
Meeting requests for the pilot will be accepted beginning 15 September 2021 and will remain open until enough have been held for the agencies to assess the program.
There are 3 stages in the PSA meeting process:
- Stage 1 – applicants request a meeting with EMA and FDA
- Stage 2 – EMA and FDA assess the meeting package, the agencies conduct a preparatory bilateral meeting, and then conduct a trilateral meeting with the applicant
- Stage 3 – EMA and FDA communicate written responses to the applicant.
During and after conclusion of the pilot, each agency will evaluate the benefits and challenges of the program, including the resources required, and determine next steps.
Read more about the general principles of the program and three stages process here.
For more details, visit the FDA website.
Contact Meditrial for immediate assistance in Europe and in the US.
We will help you to optimize your global product development program!
Table of Contents
Getting a CE mark for medical devices: a step-by-step guide
Practical guidance on the CE mark for medical devices under EU MDR, from classification and evidence to conformity assessment and EUDAMED
A Practical RBQM Framework for Medical Device Trials
Risk-based quality management for medical device trials, with practical actions for risk assessment, monitoring, data review, and trial oversight
EU Breakthrough Medical Devices Under MDCG 2025-9 (BtX)
EU breakthrough medical devices gain a clearer regulatory pathway under MDCG 2025-9, with early advice and lifecycle evidence planning
What Swissmedic Fast-Track Means for Clinical Trial Sponsors
Swissmedic swissethics Fast-Track Pilot Project accelerates eligible trial reviews and makes coordinated regulatory preparation increasingly important.
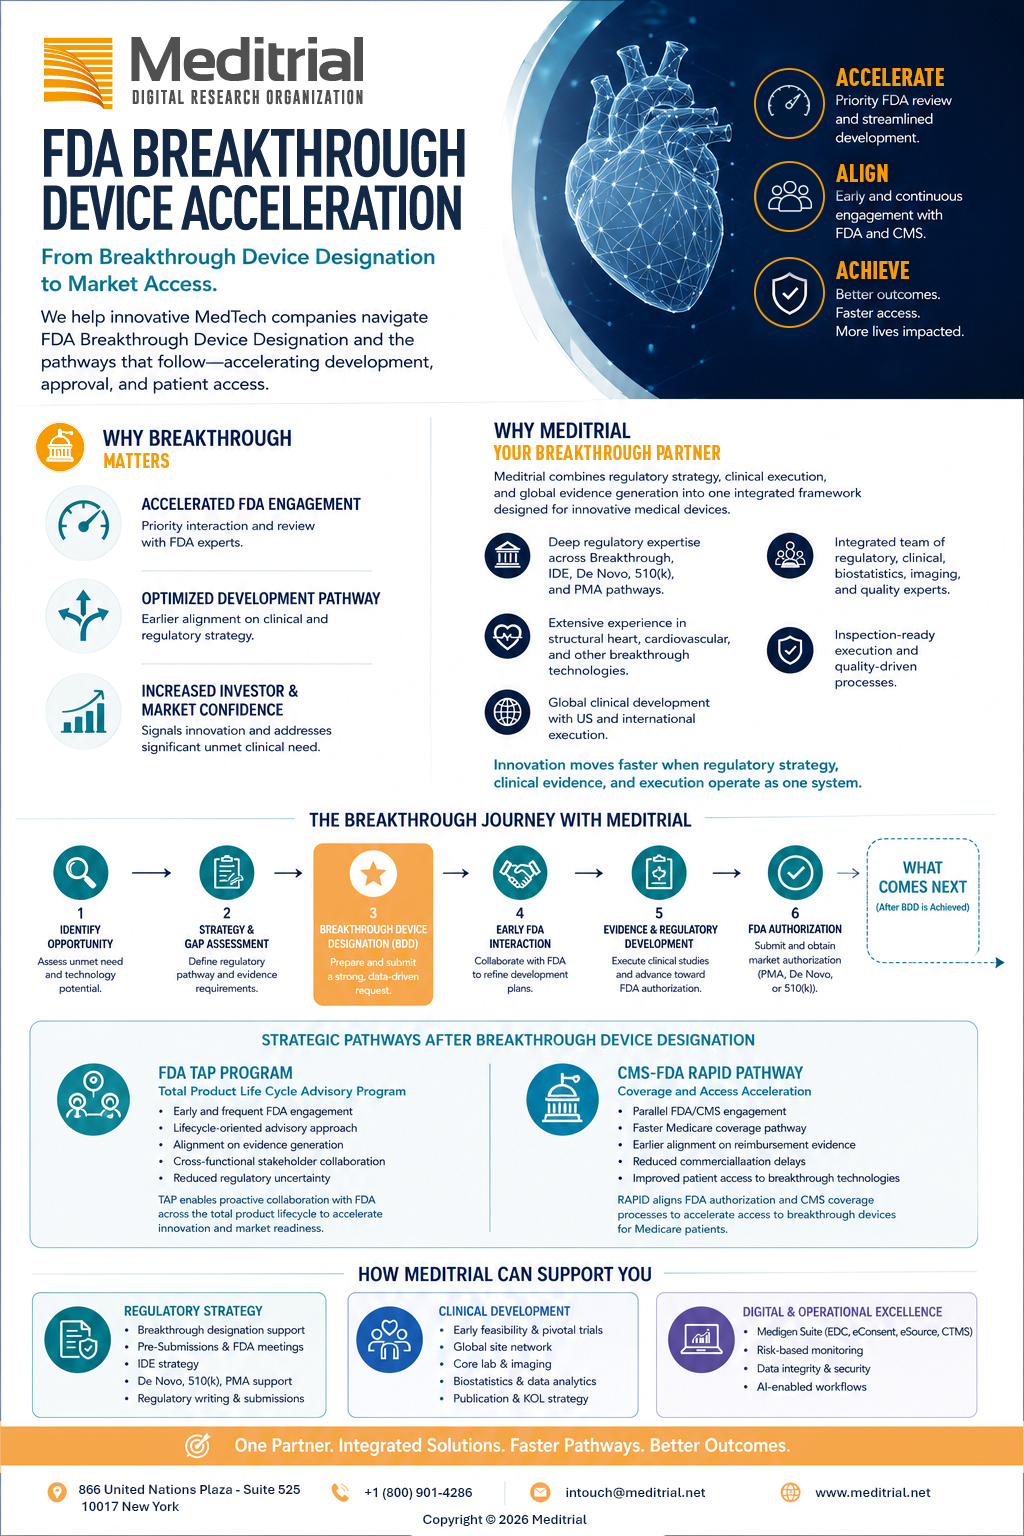
FDA Breakthrough Device Acceleration
How innovative MedTech companies can accelerate FDA approval, secure Medicare coverage, and reach patients faster—starting with Breakthrough Device Designation.
The contribution of patient associations in clinical trials
Historically, patient organizations have had the primary purpose of supporting the communities they represented. Today, this remains a crucial function, but modes have changed, as has the role that associations and patients themselves take on [...]
This article provides general information and does not constitute regulatory or legal advice. Requirements should be confirmed for the specific study, applicable regulatory framework, and markets concerned.